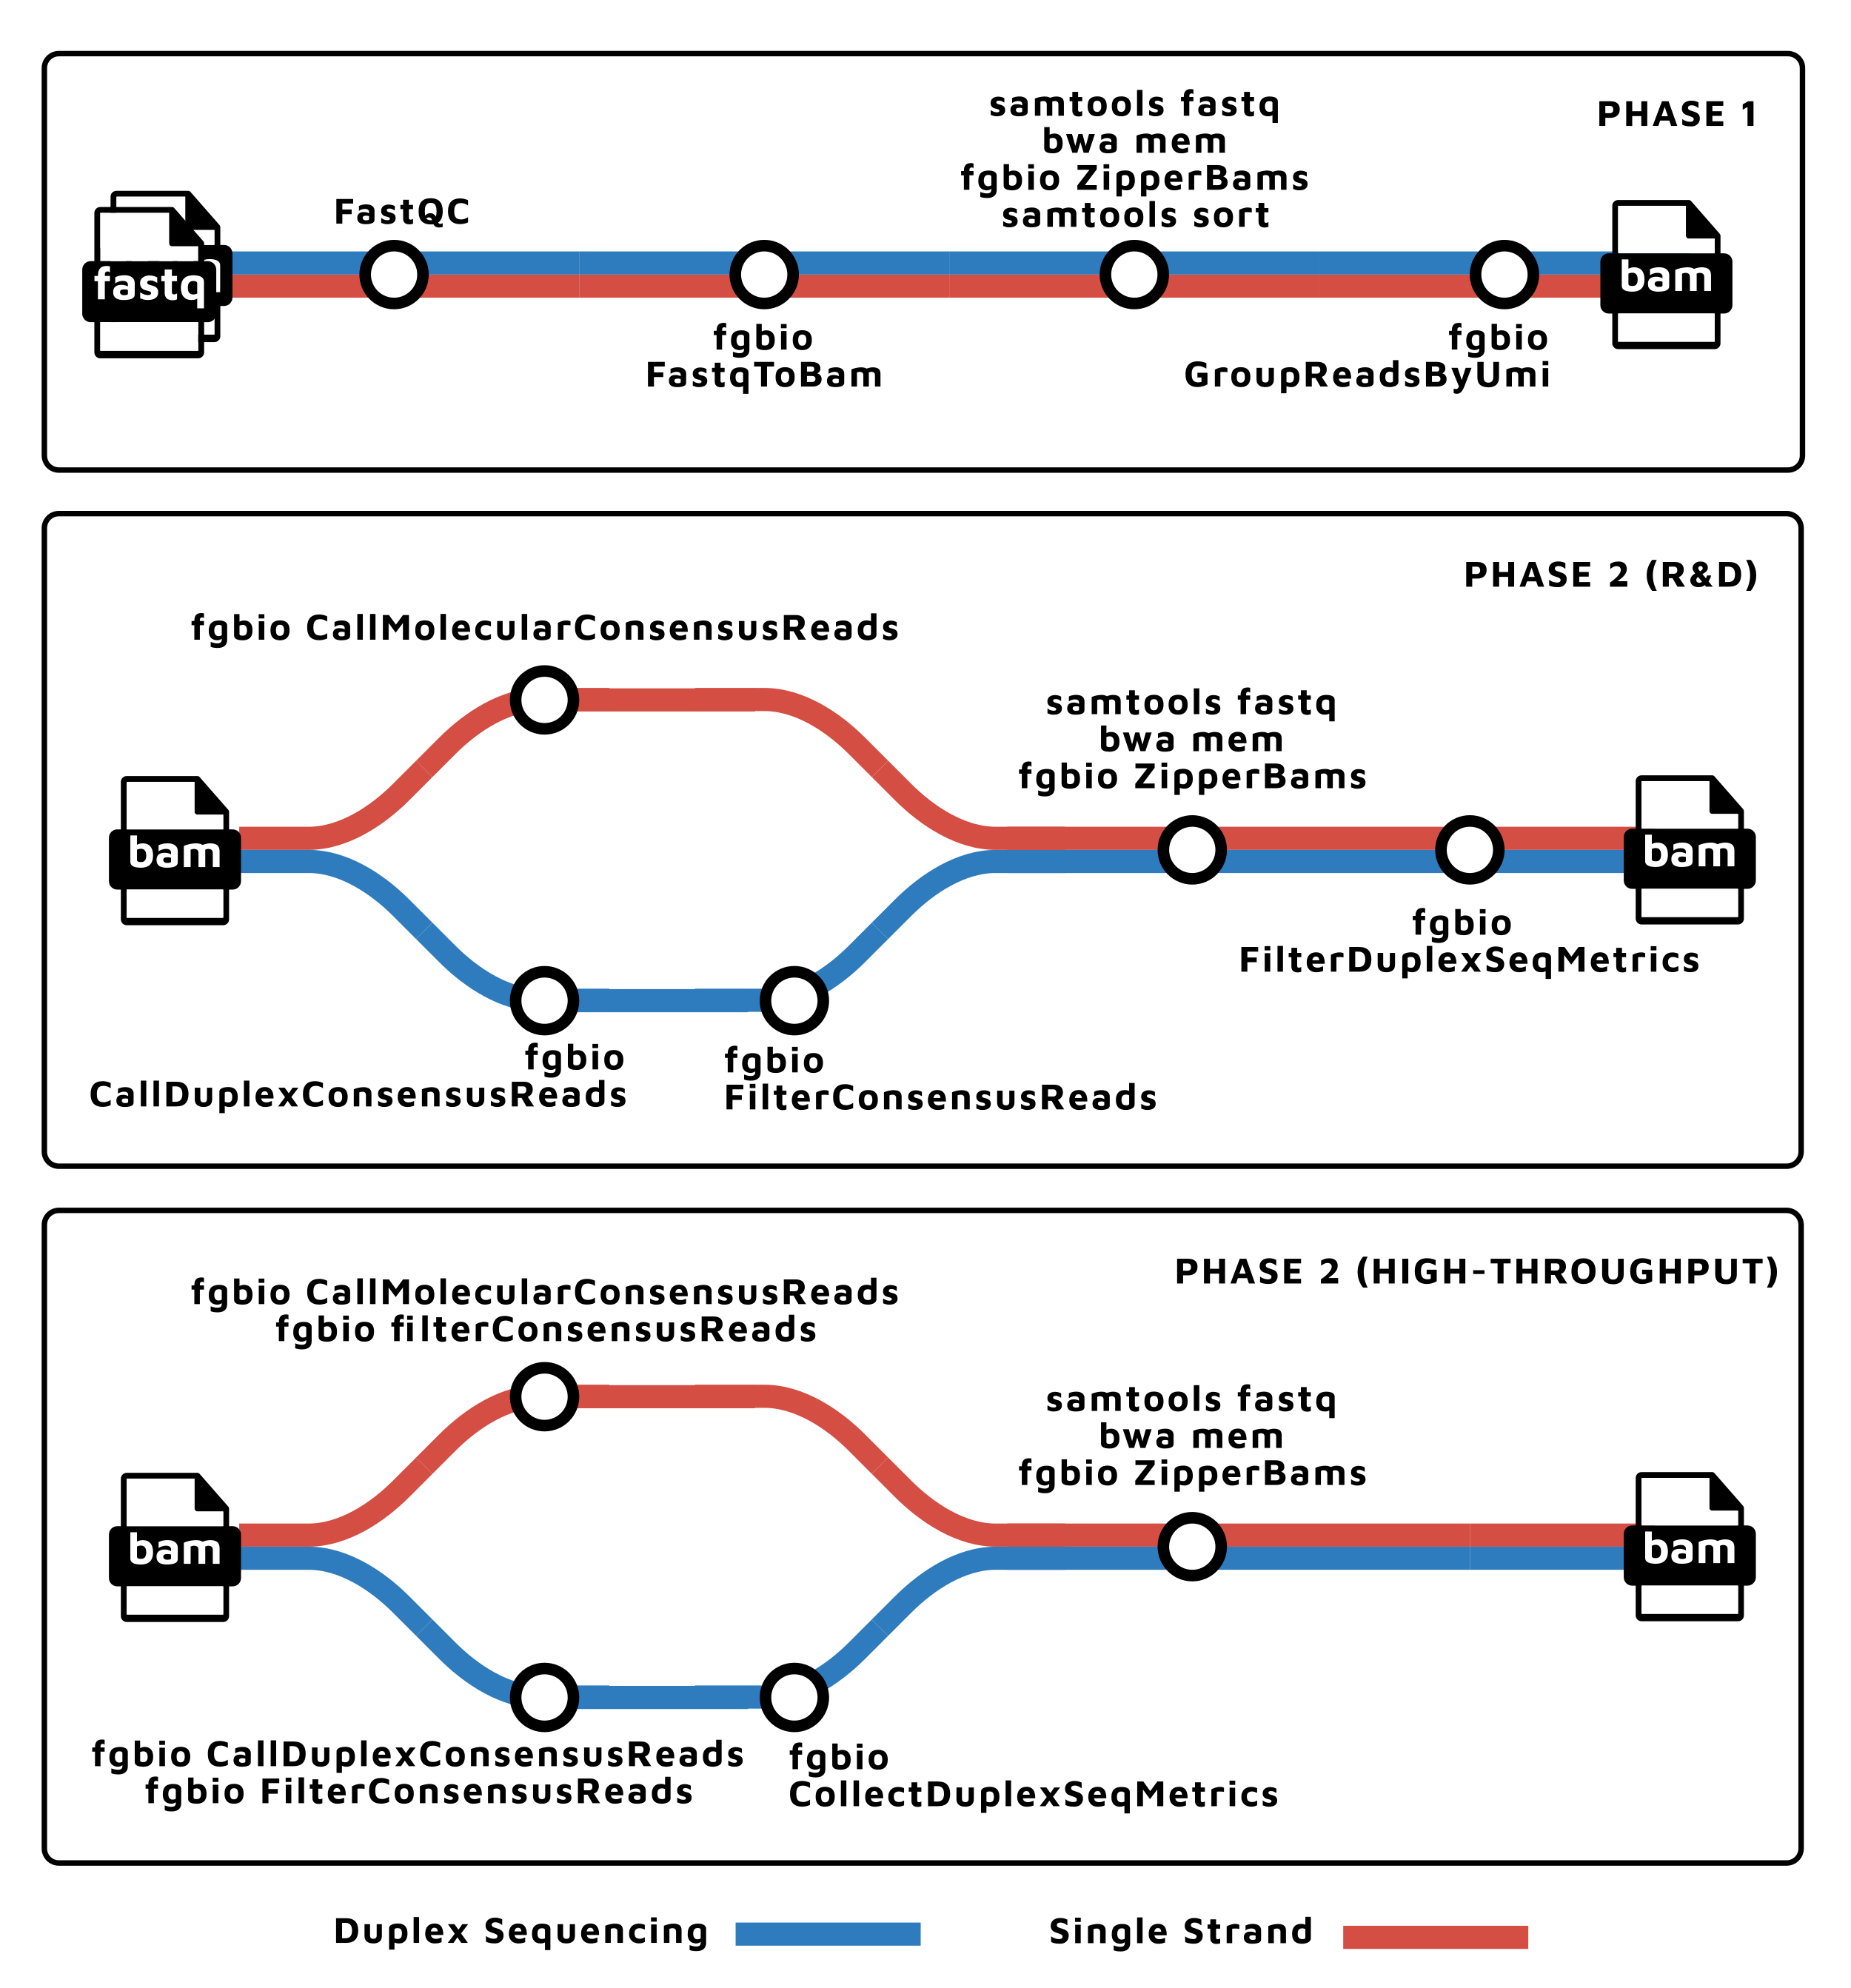
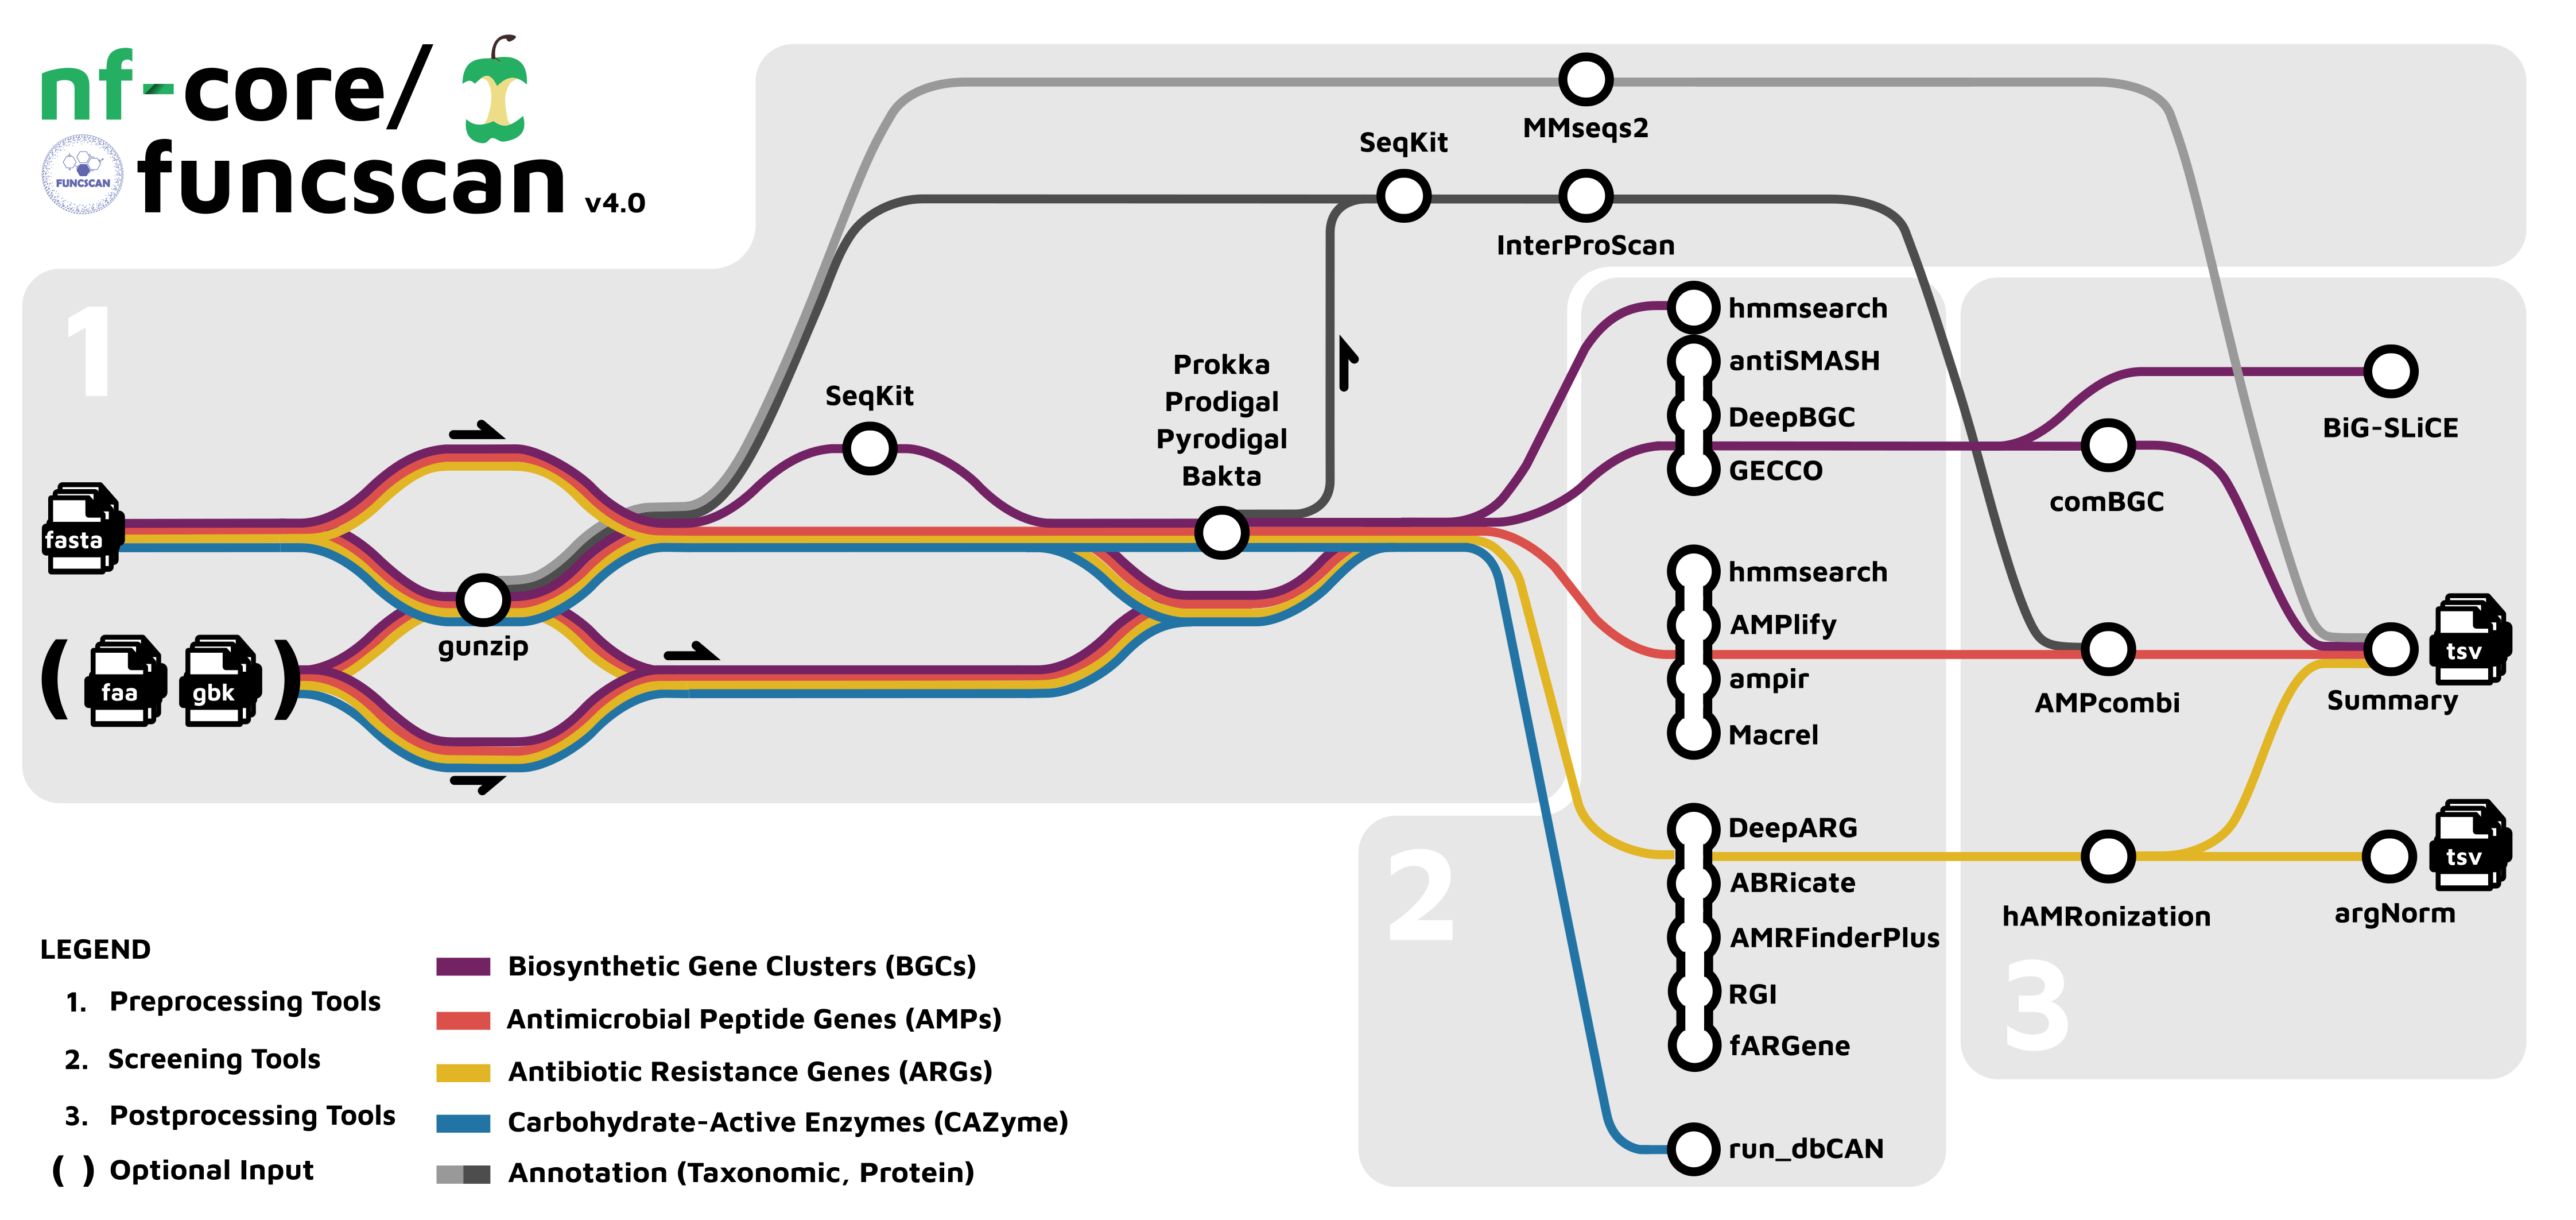
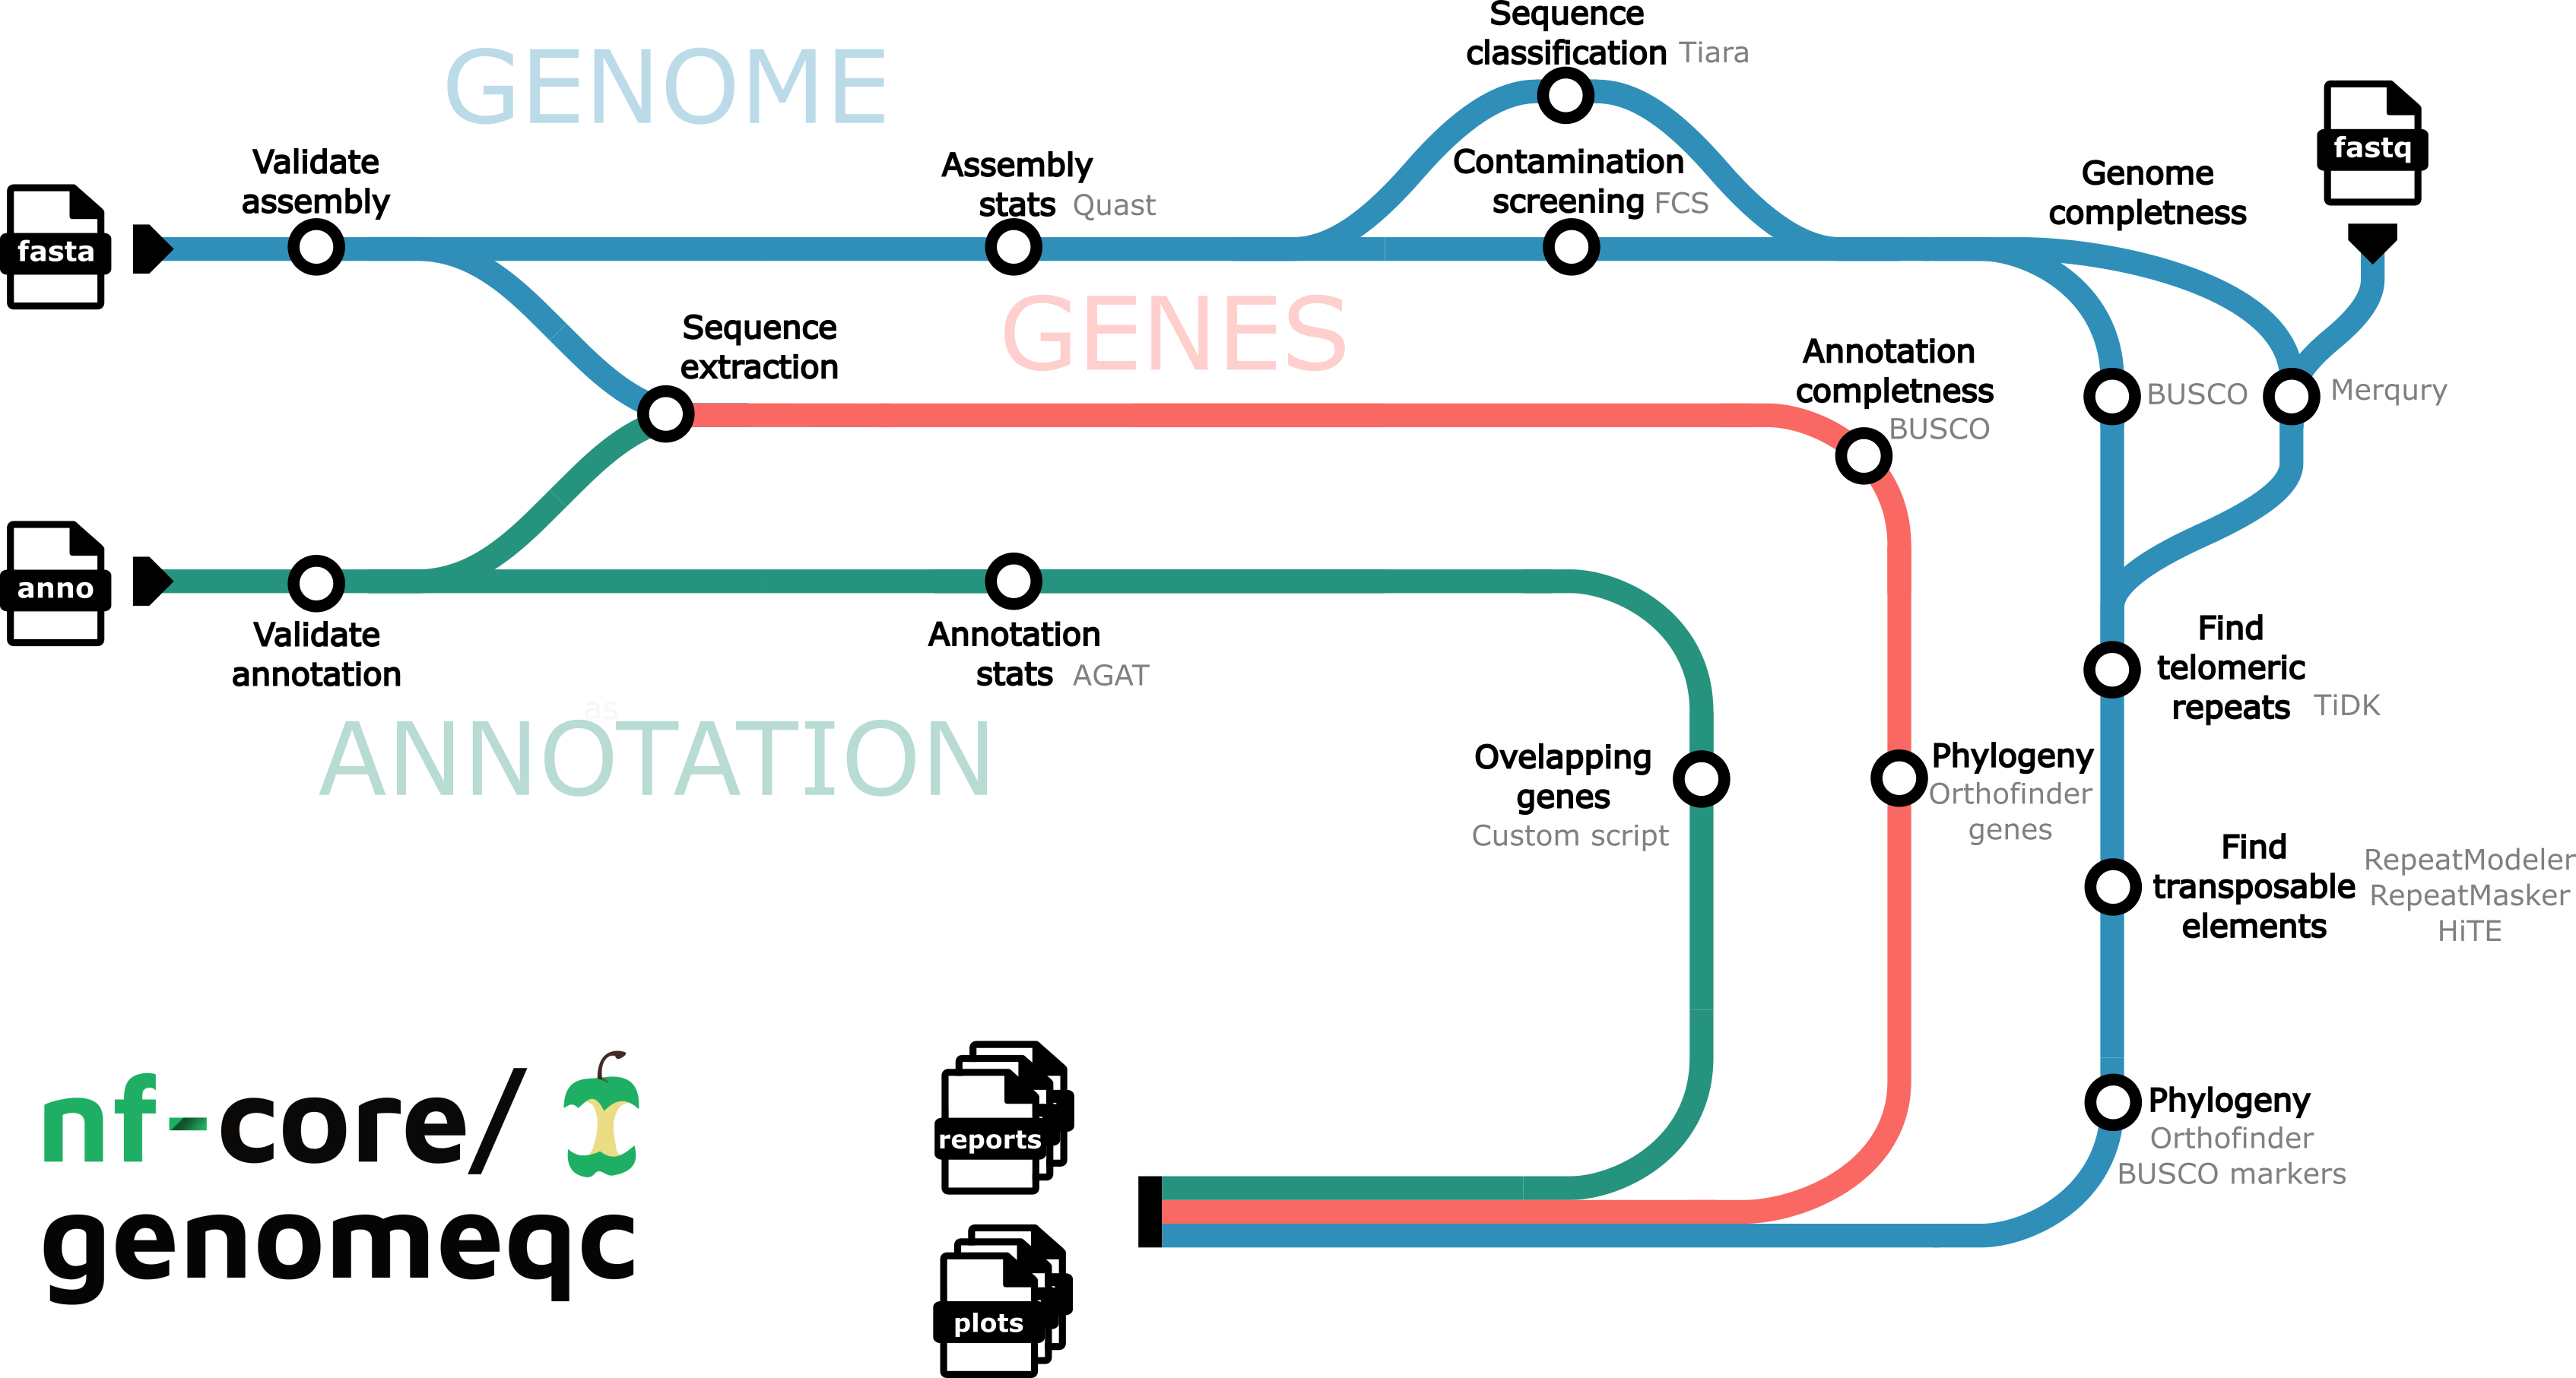
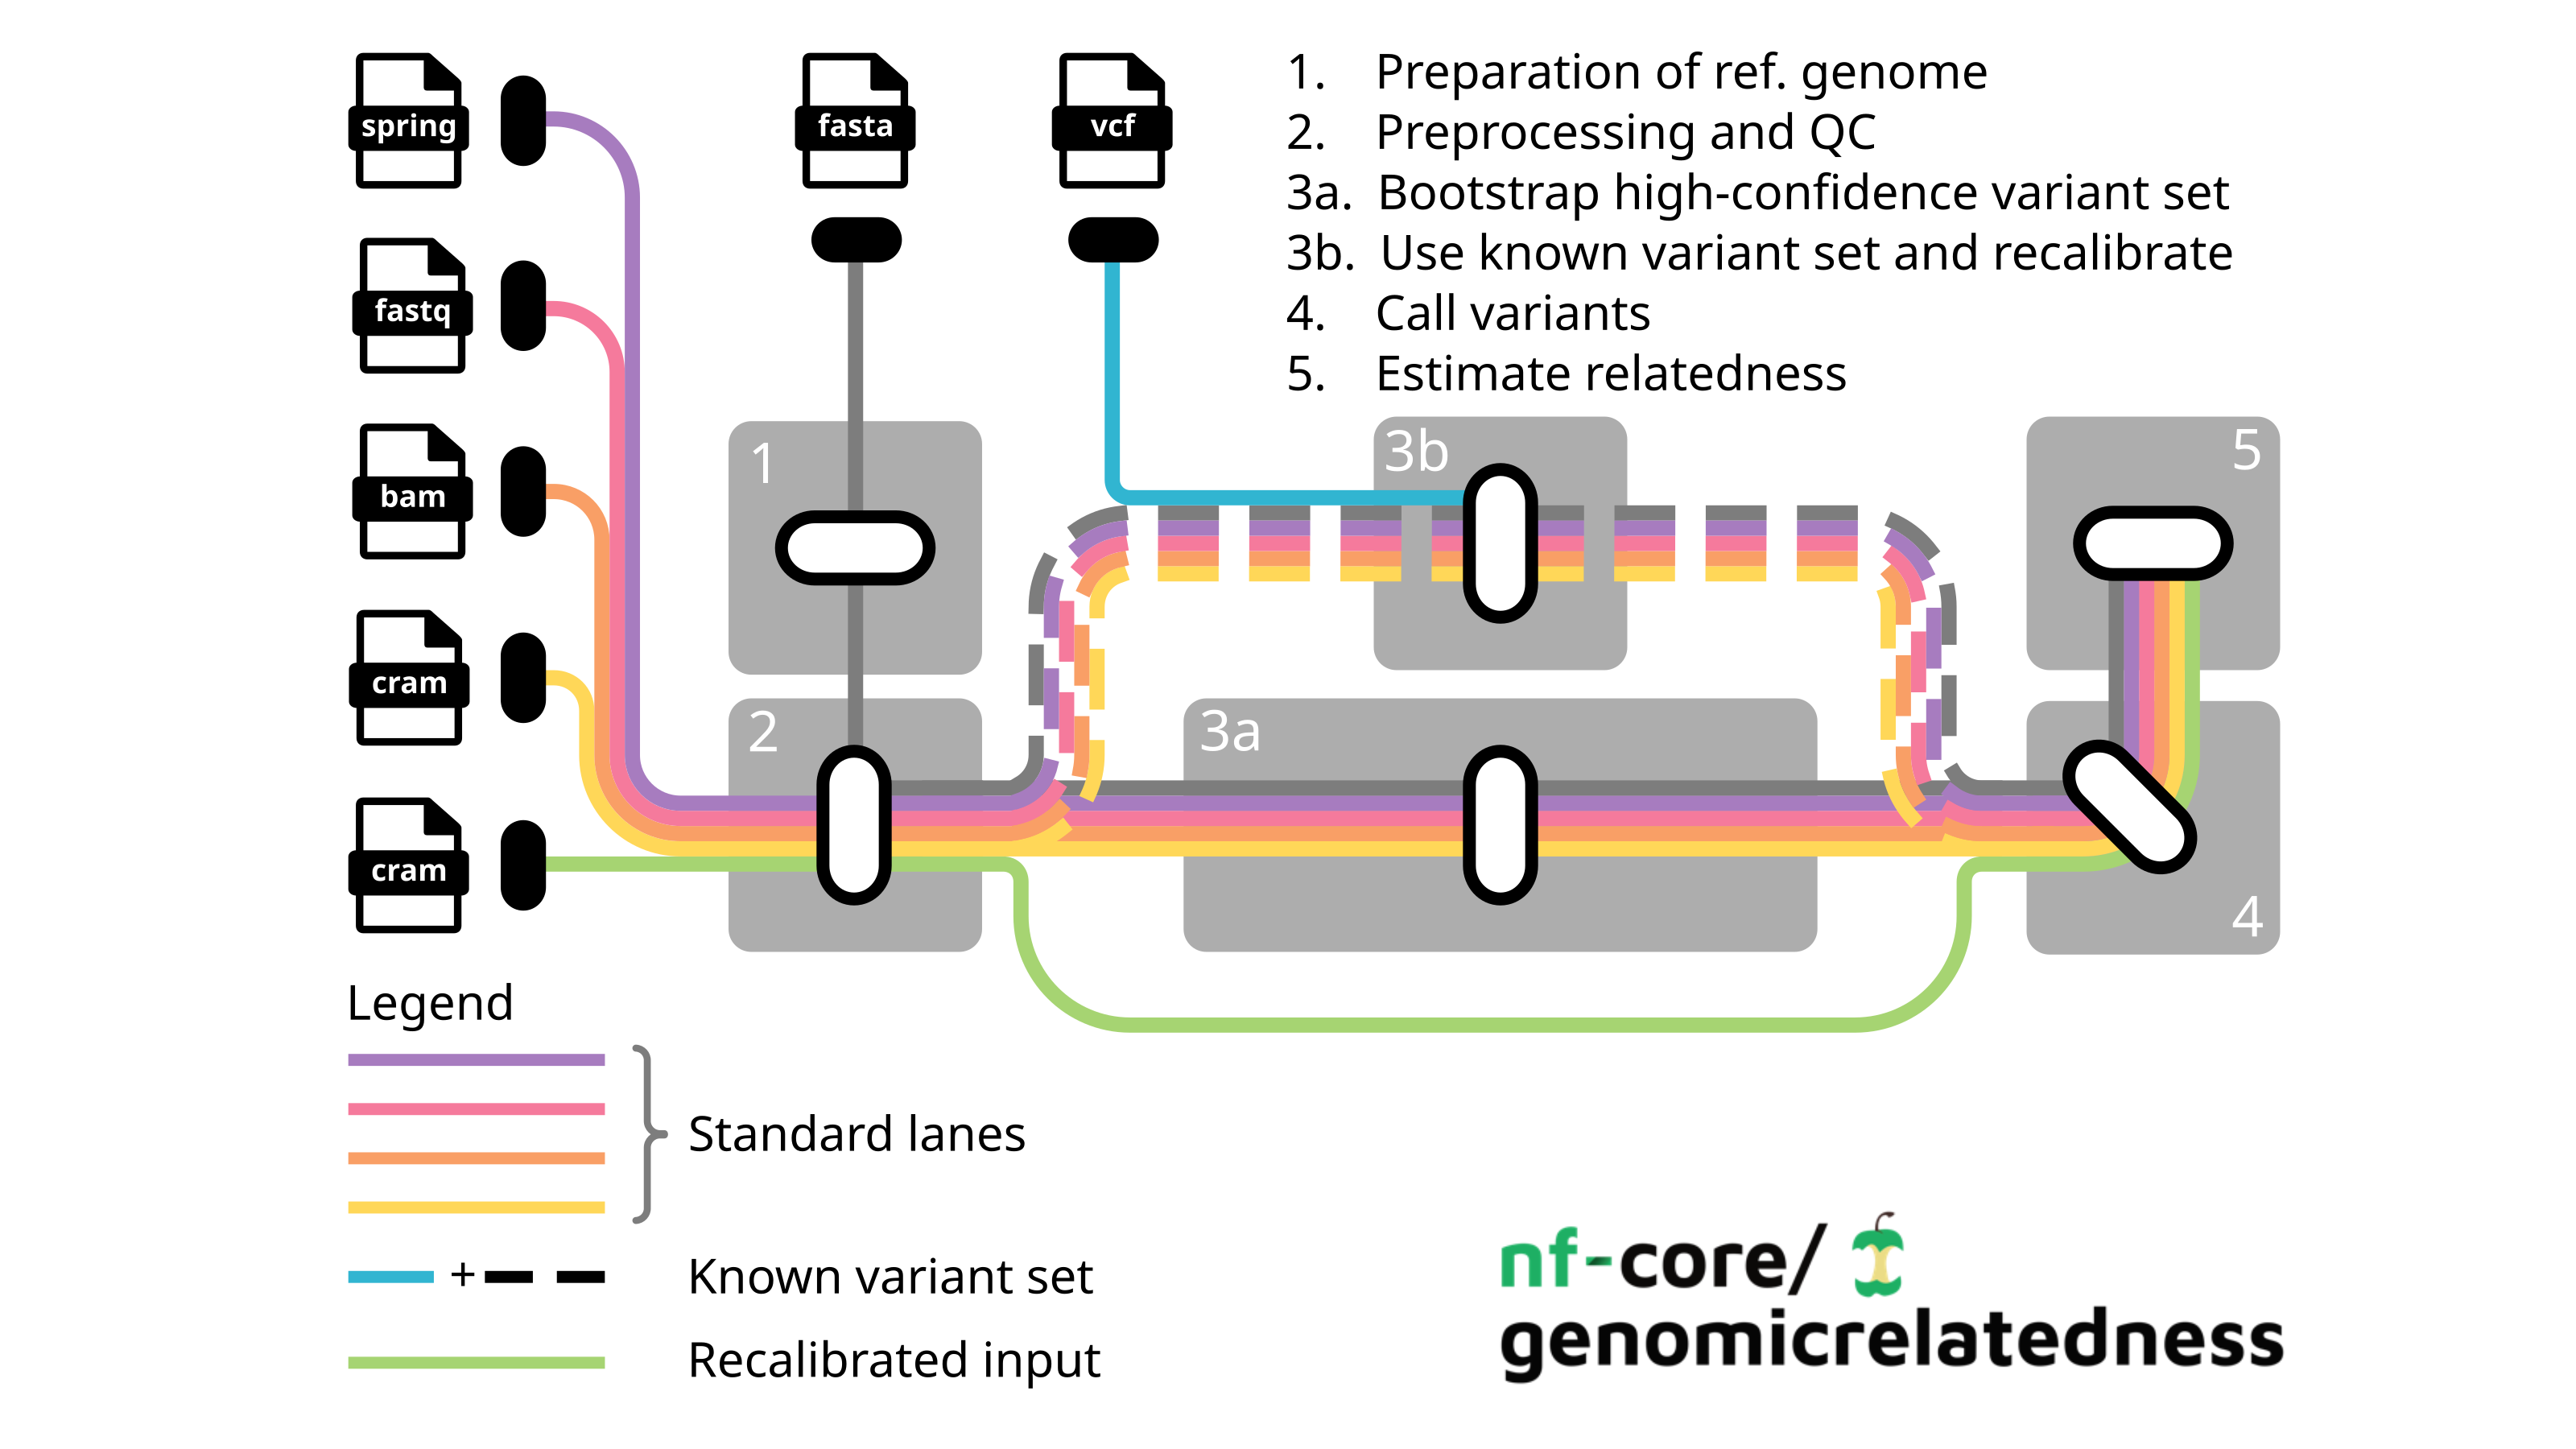
Search
Home
Pipelines
Resources
Components
Modules
Subworkflows
Configs
Shared configs
Plugins & Actions
nft-utils
nf-core-utils
setup-nextflow
setup-nf-test
Docs
Community
What's happening
Advisories
Blog
Newsletter
Events
Bytesize Talks
Hackathons
Training
Talks
Weekly Helpdesks
Programs and groups
Special interest groups
In-progress RFCs
Mentorships
Contributors
Shop
new
Connect game 👾
About
About nf-core
nf-core Lore
Stickers
Governance
Marketing
Code of conduct
Publications
Statistics
Join nf-core
Search
Select theme
Light
Dark
System
Select theme
Light
Dark
System
Join nf-core
Join
nf-core
Metro map gallery
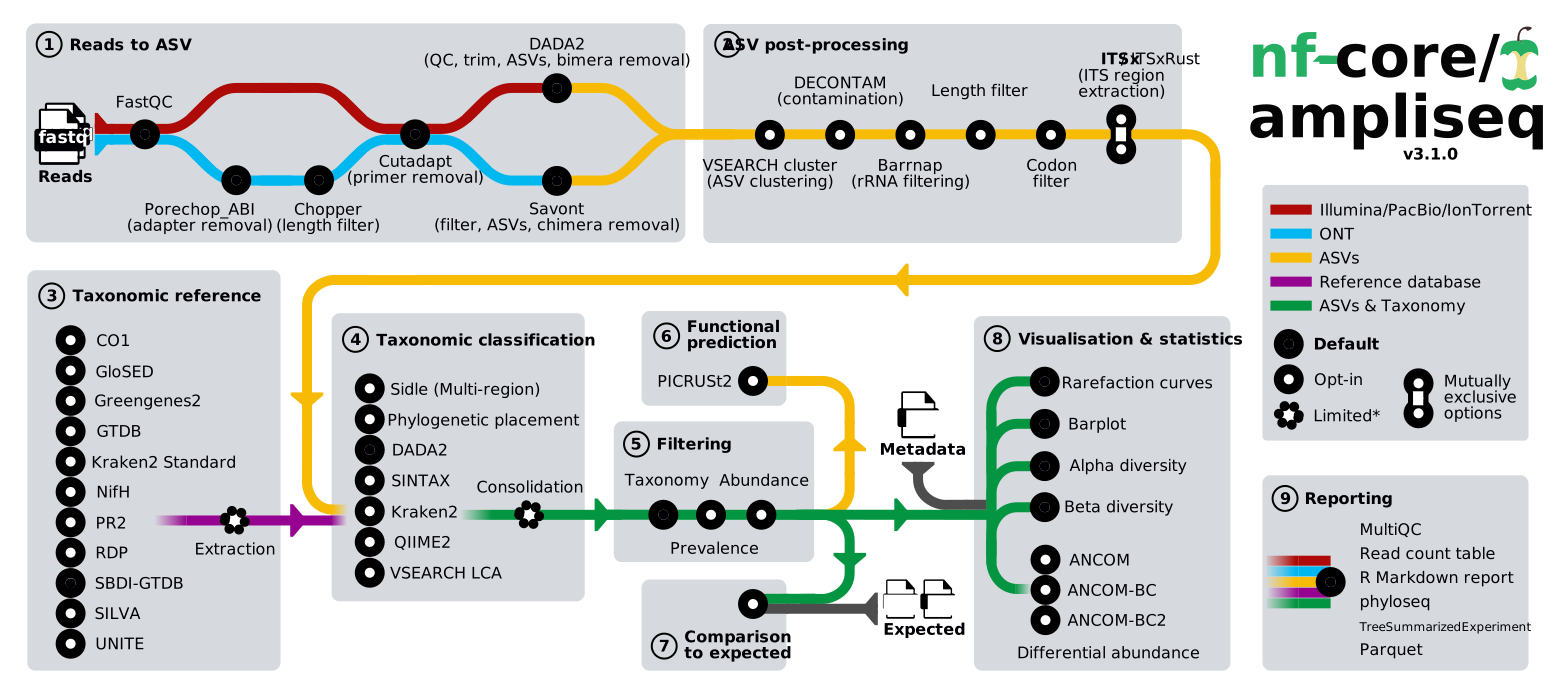
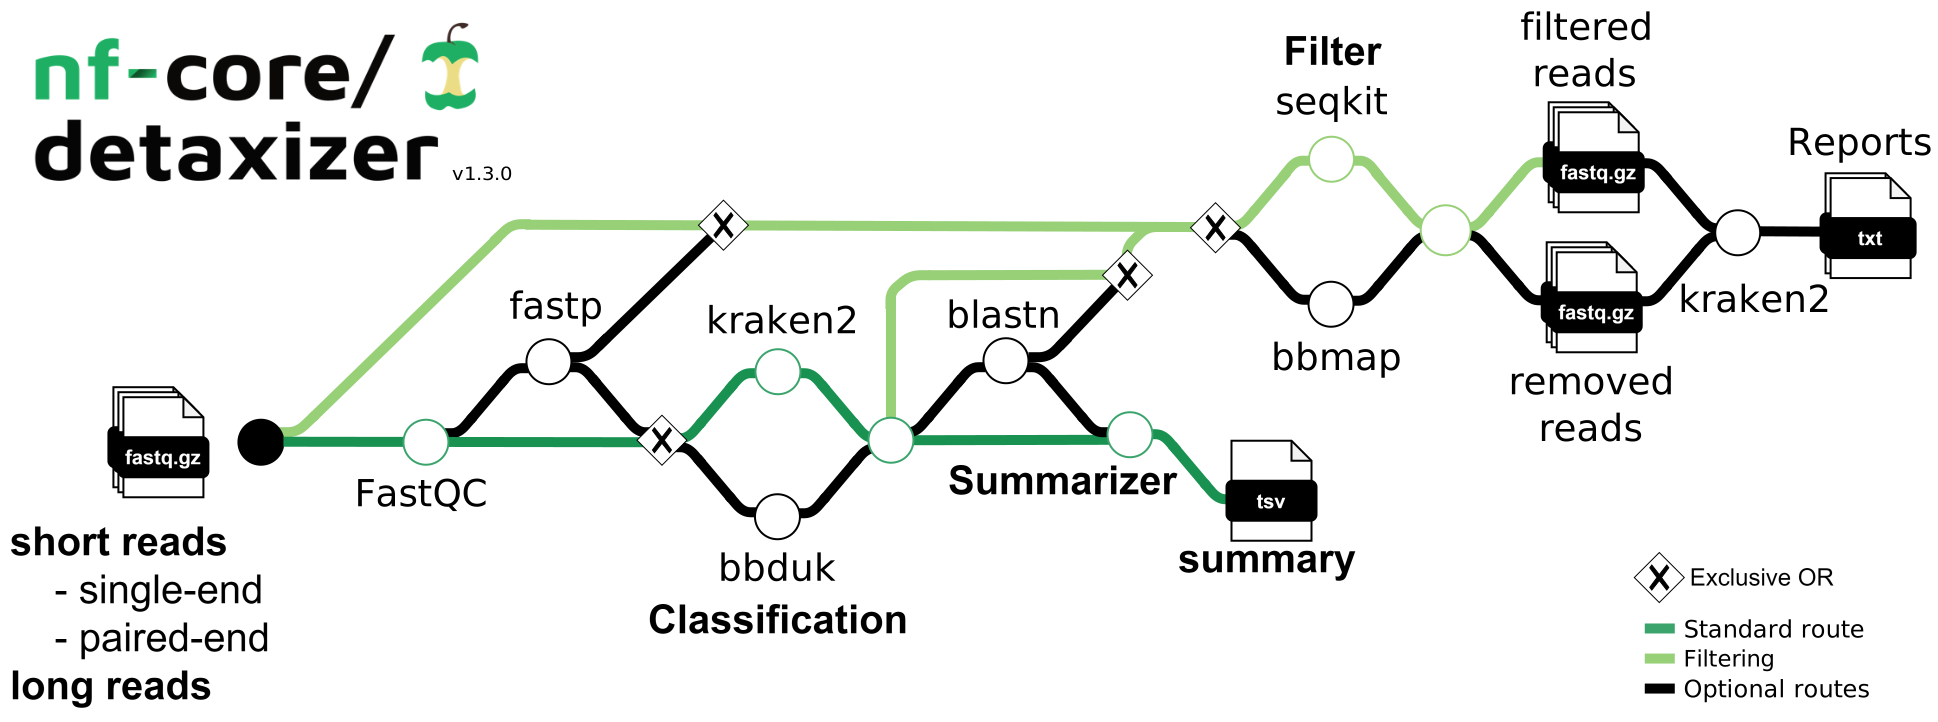
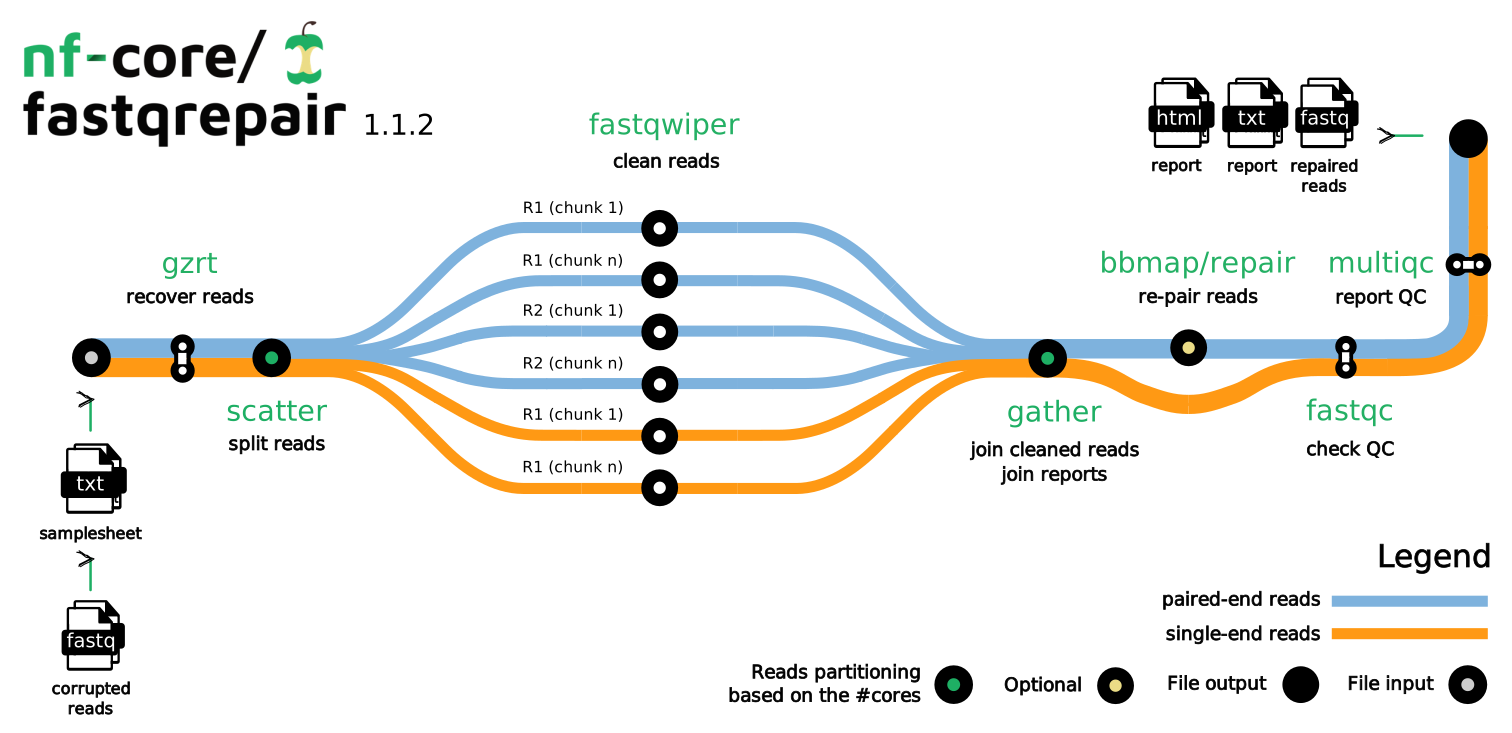
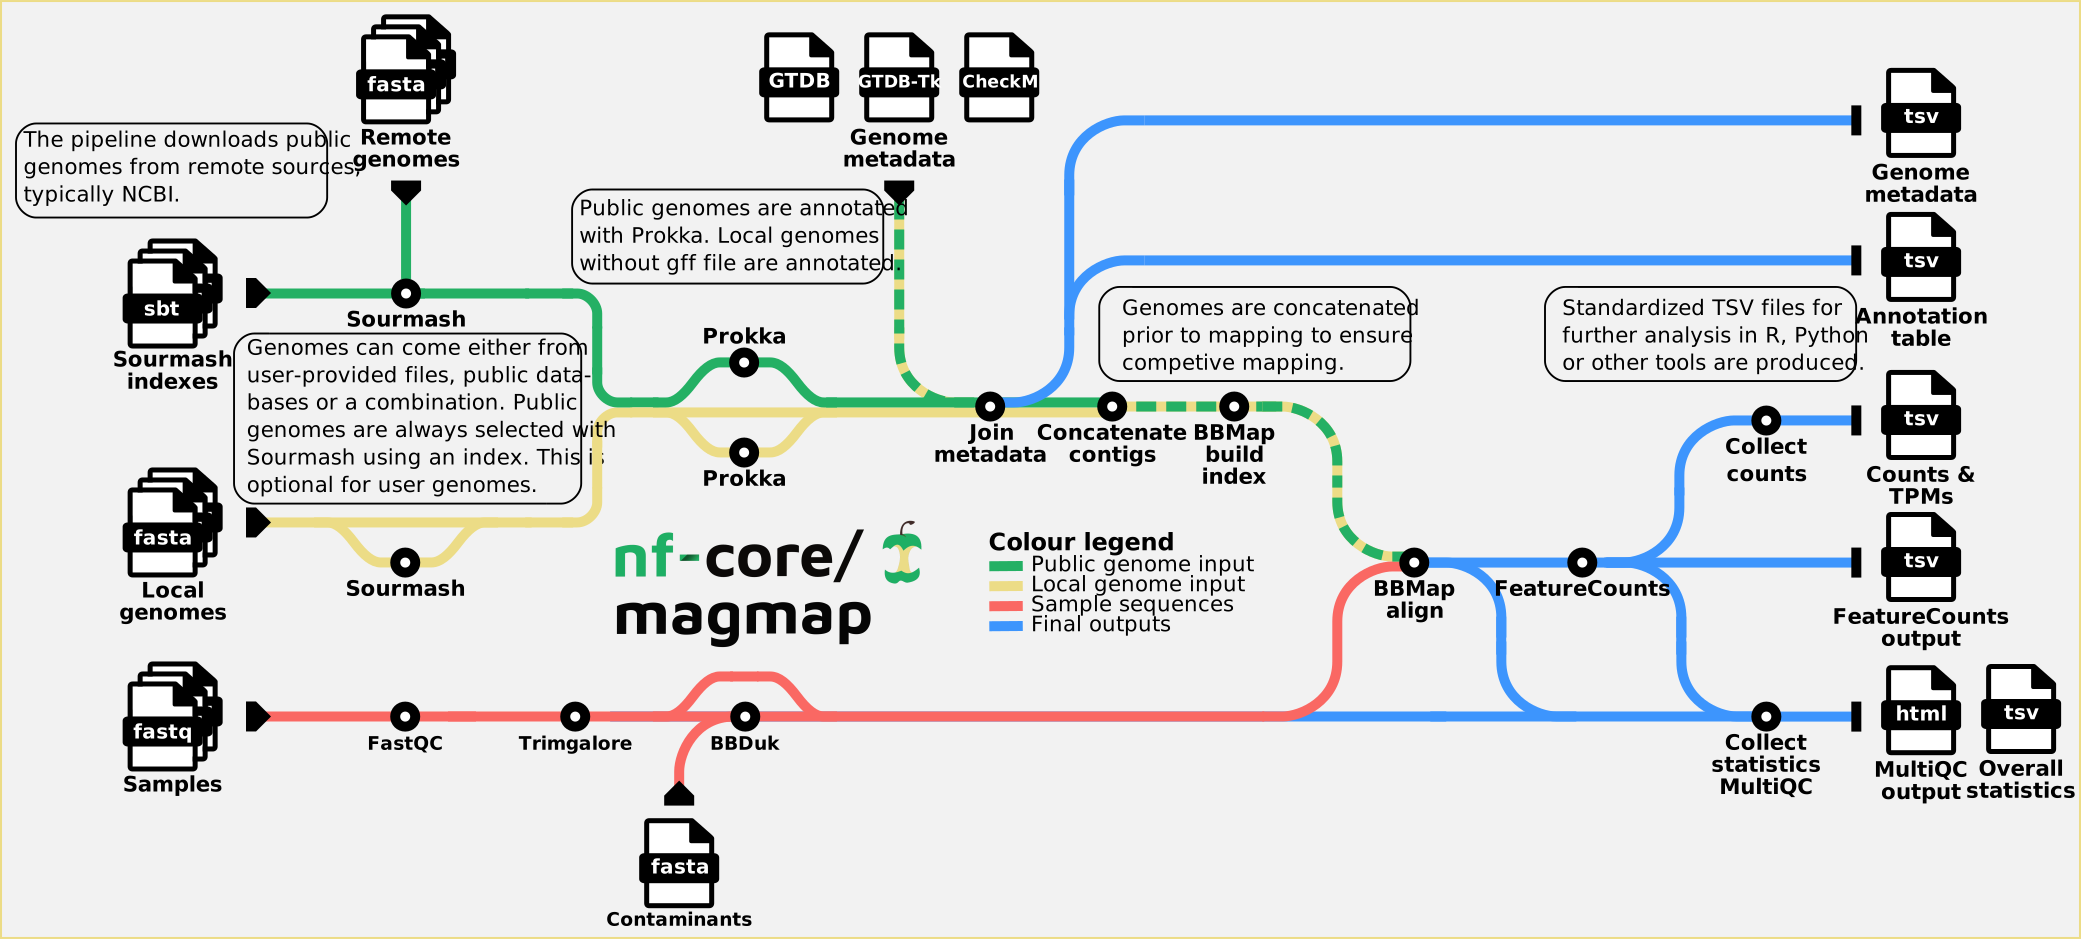
Pipeline diagrams from across nf-core. See
the docs
to find out more on how to make them.
Alpha diversity
Rarefaction curves
Differential abundance
Beta diversity
ANCOM-BC
ANCOM-BC2
Barplot
8
Visualisation & statistics
ANCOM
3
Taxonomic reference
GTDB
CO1
UNITE
GloSED
NifH
SBDI-GTDB
Greengenes2
PR2
RDP
Kraken2 Standard
SILVA
Extraction
VSEARCH cluster
(ASV clustering)
2
ASV post-processing
DECONTAM
(contamination)
Length filter
Codon
filter
(ITS region
extraction)
ITSx
/ ITSxRust
Barrnap
(rRNA filtering)
Illumina/PacBio/IonTorrent
ONT
ASVs
Reference database
ASVs & Taxonomy
Mutually
exclusive
options
Default
Opt-in
Limited*
1
Reads to ASV
Porechop_ABI
(adapter removal)
Savont
(filter, ASVs, chimera removal)
Chopper
(length filter)
FastQC
DADA2
(QC, trim, ASVs, bimera removal)
Cutadapt
(primer removal)
fastq
fastq
nf-
core/
ampliseq
Expected
Reads
Metadata
5
Taxonomy
Prevalence
Abundance
Filtering
Consolidation
4
Taxonomic classification
DADA2
QIIME2
SINTAX
Phylogenetic placement
Kraken2
VSEARCH LCA
Sidle (Multi-region)
MultiQC
Read count table
R Markdown report
phyloseq
TreeSummarizedExperiment
Parquet
9
Reporting
6
Functional
prediction
PICRUSt2
v3.1.0
7
Comparison
to expected
4) T
axonomic
and reference selection
4) Taxonomic and reference selection
Short Reads
Short Reads
Long Reads
Long Reads
Hybrid Sequencing
Hybrid Sequencing
Multiple
parallel
options
Multiple parallel op...
Single choice
Single choice
Default
Default
Opt-in
Opt-in
MULTIQC
MULTI...
Fasta
Fasta
Pipeline input
Pipeline input
Pipeline output
Pipeline output
v2.7
v2.7
3) Polishing
3) Polishing
Fast5
Fast5
FastQC
FastQC
FastP
FastP
Fastq
Fastq
Fastq
Fastq
Toulligqc
Nanoplot
Toulligqc...
Filtlong
Porechop
Filtlong...
1) Pre-porcessing and Quality Control
1) Pre-porcessing and Quality Control
cat FastQ
cat Fas...
cat FastQ
cat FastQ
Megahit
Megahit
5) Genome annotation
5) Genome annotation
Autocycler
Miniasm
Flye
Canu
Raven
Autocycle...
2) Genome assembly
2) Genome assembly
Unicycler
Unicycler
Medaka
Medaka
Nanopolish
Nanopolish
Kraken2
Kraken2
Kmerfinder
Kmerfinder
Rasusa
Rasusa
DragonFlye
DragonFlye
6)
Assembly QC
&
reporting
6) Assembly QC &...
QUAST
BUSCO
QUAST...
MultiQC
MultiQC
Dfast
Liftoff
Bakta
Prokka
Dfast...
v1.3.0
fastp
FastQC
kraken2
bbduk
blastn
seqkit
bbmap
filtered
reads
kraken2
removed
reads
Reports
Filtering
Standard route
Optional routes
1.1.2
gzrt
scatter
gather
fastqwiper
bbmap/repair
fastqc
multiqc
R1 (chunk 1)
R1 (chunk 1)
R1 (chunk n)
R1 (chunk
n
)
R2 (chunk 1)
R2 (chunk
n
)
recover reads
split reads
join cleaned reads
join reports
clean reads
re-pair reads
check QC
report QC
Legend
paired-end reads
File input
File output
Optional
Reads partitioning
based on the #cores
single-end reads
txt
report
txt
report
txt
report
html
report
fastq
repaired
reads
txt
report
txt
samplesheet
fastq
corrupted
reads
1
Download metadata
2
Download FASTQ files
3
Downstream
CSV
CSV
FASTQ
CSV
nf-core/<PIPELINE>
Aspera
Aspera
fastq-dl
fastq-dl
FTP
FTP
sra-tools
sra-tools
Fetch metadata
Fetch metadata
Get
download
links
Get
download
links
SRA
GEO
ENA
DDBJ
created with nf-metro v2.1.0
License: CC BY 4.0
nf-core/genomeassembler
1
Input
2
Assemble
3
Polish
4
Scaffold
5
QC
6
Annotation Liftover
FASTQ
ONT reads
FASTQ
HiFi reads
FASTA
Assembly
FASTA
Assembly
FASTQ
ONT reads (trimmed)
FASTQ
short reads
FASTA
polished Assembly
FASTA
Reference
FASTA
(polished) Assembly
FASTQ
ONT reads
FASTQ
HiC reads
FASTA
Scaffolded assembly
FASTA
Assembly
FASTQ
short reads (trimmed)
FASTA
Reference
HTML
Report
FASTA
Assembly
FASTA
Reference
GFF
Reference
GFF
Lifted annotations
fastplong
fastplong
fastplong
fastplong
hifiasm /
flye
hifiasm /
flye
hifiasm --ul
hifiasm --ul
ONT assembly
ONT assembly
HiFi assembly
HiFi assembly
medaka / dorado
medaka / dorado
fastp
fastp
LINKS
longstich
LINKS
longstich
RagTag
RagTag
BUSCO
BUSCO
fastp
fastp
QUAST
QUAST
meryl
meryl
liftoff
liftoff
RagTag
Scaffold
RagTag
Scaffold
pilon
pilon
yahs
yahs
merqury
merqury
Assemble: Single ONT
Assemble: Single HiFi
Assemble: Hybrid
Assemble: Scaffold
Polish: pilon
Polish: ONT
Scaffold: Long reads
Scaffold: Reference
Scaffold: HiC
QC
Annotation Liftover
created with nf-metro v1.1.0
Sourmash
Prokka
Prokka
fastq
fastq
fasta
Local
genomes
fastq
fastq
sbt
Sourmash
indexes
fastq
fastq
fastq
Samples
FastQC
Join
metadata
GTDB
GTDB-Tk
CheckM
Genome
metadata
Concatenate
contigs
BBMap
build
index
BBMap
align
FeatureCounts
Collect
statistics
MultiQC
fastq
fastq
fasta
Remote
genomes
Sourmash
Collect
counts
html
MultiQC
output
tsv
Overall
statistics
tsv
FeatureCounts
output
tsv
Counts &
TPMs
tsv
Genome
metadata
tsv
Annotation
table
Public genome input
Local genome input
Sample sequences
Final outputs
Colour legend
Genomes can come either from
user-provided files, public data-
bases or a combination. Public
genomes are always selected with
Sourmash using an index. This is
optional for user genomes.
Standardized TSV files for
further analysis in R, Python
or other tools are produced.
Genomes are concatenated
prior to mapping to ensure
competive mapping.
Public genomes are annotated
with Prokka. Local genomes
without gff file are annotated.
The pipeline downloads public
genomes from remote sources,
typically NCBI.
Trimgalore
BBDuk
fasta
Contaminants
csv
csv
csv
csv
metaboigniter
metaboigniter
/
/
pre-processing
pre-processing
Sirius & CSI:FingerID
Sirius & CSI:FingerI...
input files
input files
samplesheet check
samplesheet check
peak picking
peak picking
feature detection
feature detection
map alignment
map alignment
adduct detection
adduct detection
feature linking
feature linking
structure prediction
structure prediction
analogue search
analogue search
annotation
annotation
re-quantification
re-quantification
csv
csv
csv
csv
mzML
mzML
samplesheet
samplesheet
raw LC-MS data
raw LC-MS data
Feature Matrix
Feature Matrix
mandatory processes
mandatory processes
optional processes
optional processes
re-quantification
re-quantification
formula prediction
formula prediction
MS2Query
MS2Query
Text is not SVG - cannot display
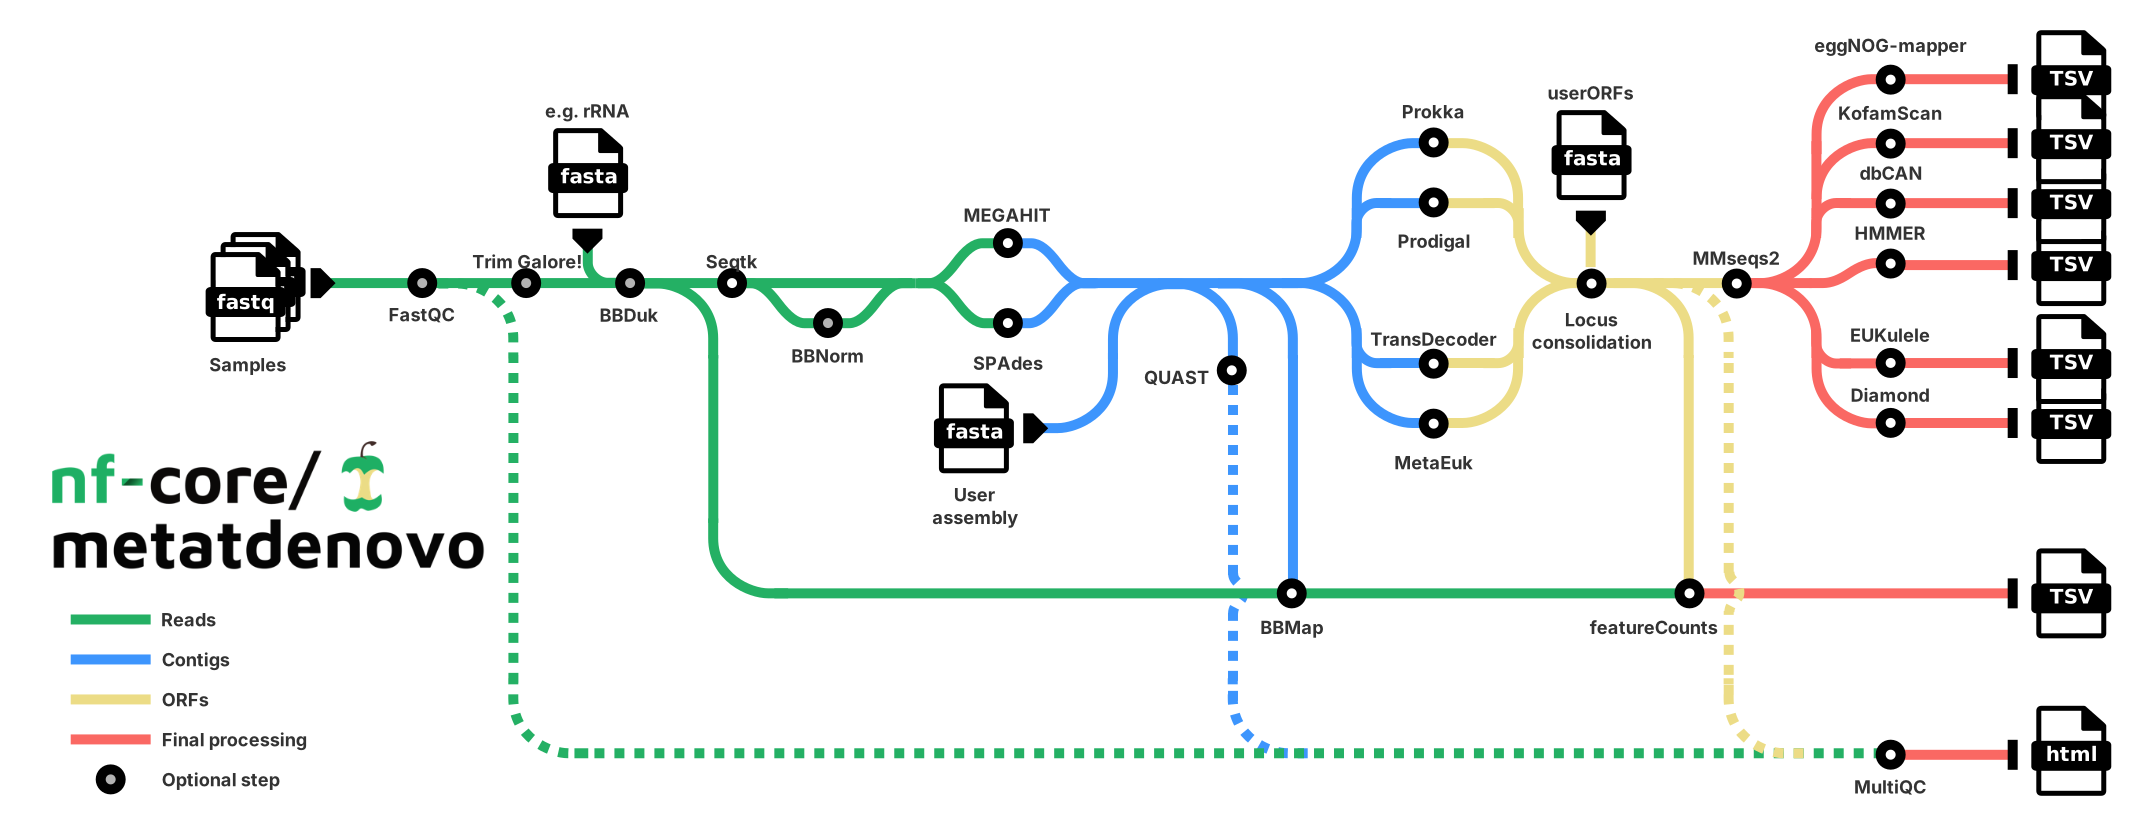
FastQC
Trim Galore!
fastq
fastq
fastq
fasta
BBDuk
Seqtk
BBNorm
MEGAHIT
SPAdes
fasta
Samples
e.g. rRNA
User
assembly
QUAST
Prokka
Prodigal
TransDecoder
MetaEuk
BBMap
Locus
consolidation
MMseqs2
featureCounts
eggNOG-mapper
KofamScan
dbCAN
HMMER
EUKulele
TSV
Diamond
fasta
userORFs
TSV
TSV
TSV
TSV
TSV
html
MultiQC
TSV
Reads
Contigs
ORFs
Final processing
Optional step
Bespoke Script
Compile report
.fastq
Plasmodium reads
fastqc
bbduk
.fasta
Plasmodium
and Anopheles references
minimap2
clair3
TBD
GATK
BWA-MEM3
hmmibd-rs
Quality control
Remove vector reads
Align samples
MOI/Compositional haplotype calling
Relatedness analysis
.bam
.vcf
Short reads
Long reads
Reference genomes
Legend
IQTree
Phylogenetic tree
Space for expanding population-aware layer
nf-core/
pathogenepidemiology
v0.1.0dev
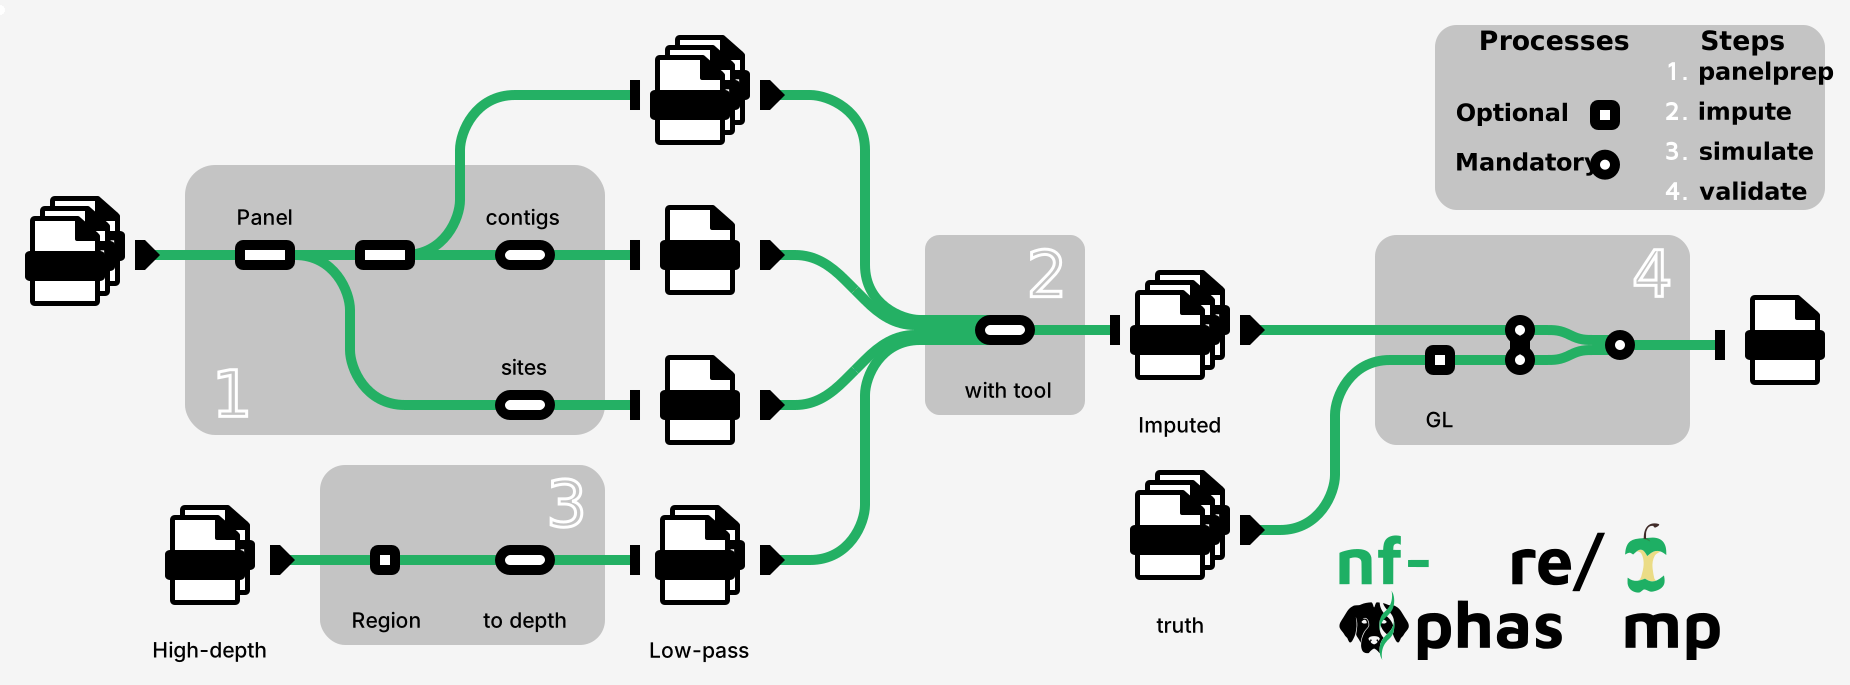
Steps
Processes
panelprep
simulate
validate
Optional
Mandatory
impute
Normalize
Panel
Phase
Chunk
contigs
Extract
sites
Impute
with tool
Statistics
Compute
GL
Concordance
Downsample
to depth
Extract
Region
Sample
Low-pass
Chunks
Panel
Sites
Panel
Sample
High-depth
Sample
truth
Sample
Imputed
Statistics
1
2
3
4
txt
tsv
fastq
bam
fastq
bam
fastq
fastq
vcf
fastq
fastq
vcf
fastq
fastq
vcf
fastq
fastq
vcf/bam
csv
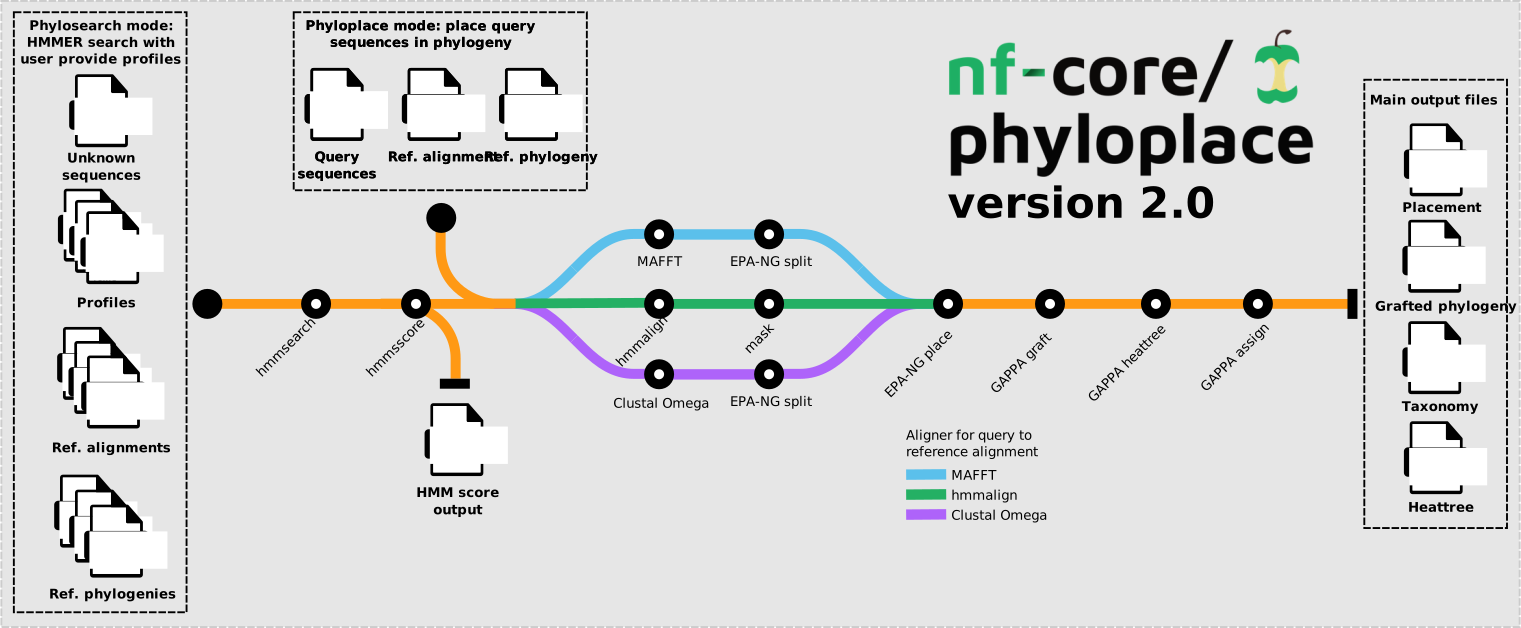
hmmalign
mask
EPA-NG place
GAPPA graft
GAPPA heattree
GAPPA assign
MAFFT
EPA-NG split
hmmsearch
hmmsscore
HMM score
output
Clustal Omega
EPA-NG split
Phylosearch mode:
HMMER search with
user provide profiles
Unknown
sequences
Profile
Profile
Profiles
Ref. alignments
Ref. phylogenies
Query
sequences
Ref. phylogeny
Ref. alignment
Phyloplace mode: place query
sequences in phylogeny
Query
sequences
Ref. phylogeny
Ref. alignment
Phyloplace mode: place query
sequences in phylogeny
Main output files
Grafted phylogeny
Taxonomy
Heattree
Placement
Clustal Omega
hmmalign
MAFFT
Aligner for query to
reference alignment
version 2.0
4
4
1
1
2
2
STAGE
1. Check quality and pre-process
2. Annotate domains
3. Annotate functions
4. Predict secondary structures
STAGE...
seqfu
stats
seqfu...
seqfu
stats
seqfu...
seqkit
seq
seqkit...
seqkit
replace
seqkit...
seqkit
rmdup
seqkit...
FAA
FAA
Amino acid sequence annotation
Database download
Amino acid sequence annotation...
METHOD
METHOD
s4pred
runmodel
s4pred...
HTML
HTML
MultiQC
MultiQC
aria2
aria2
PFAM
PFAM
FUNFAM
FUNFAM
hmmer
hmmsearch
hmmer...
DOMTBL
DOMTBL
SS2
SS2
3
3
interproscan
interproscan
GFF3
GFF3
TSV
TSV
JSON
JSON
XML
XML
1
2
3
4
5
6
7
seqfu
stats
seqkit
seq
seqkit
replace
seqkit
rmdup
seqfu
stats
gunzip
untar
find
concatenate
untar
hmmer
hmmsearch
mmseqs
createdb
mmseqs
cluster
mmseqs
linclust
mmseqs
createtsv
gunzip
hmmer
eslsfetchindex
mgnifam
generatefamilies
hhsuite
reformat
seqkit
seq
famsa
align
mafft
align
clipkit
hmmer
hmmbuild
hmmer
hmmsearch
hmmer
hmmalign
find
concatenate
hmmer
hmmsearch
mmseqs
createdb
mmseqs
linclust
mmseqs
cluster
mmseqs
createtsv
famsa
align
mafft
align
hhsuite
reformat
tar
find
concatenate
cmaple
multiqc
FAA
sequences
MSA
full MSAs
HMM
HMMs
MSA
seed MSAs
HMM
HMM library
MSA
seed MSAs
MSA
full MSAs
FAA
sequences
TREEFILE
HTML
MultiQC
CSV
proteinfold /
proteinannotator
Create new protein families
(Optional) Update existing families
(Optional) Merge similar families
Runs per sample
Runs per family
STAGE
1. Check quality and pre-process
2. (Optional) Search existing families
3. Cluster sequences
4. Generate family models & MSAs
5. (Optional) Remove redundancies
6. (Optional) Build phylogenetic trees
7. Create downstream samplesheet
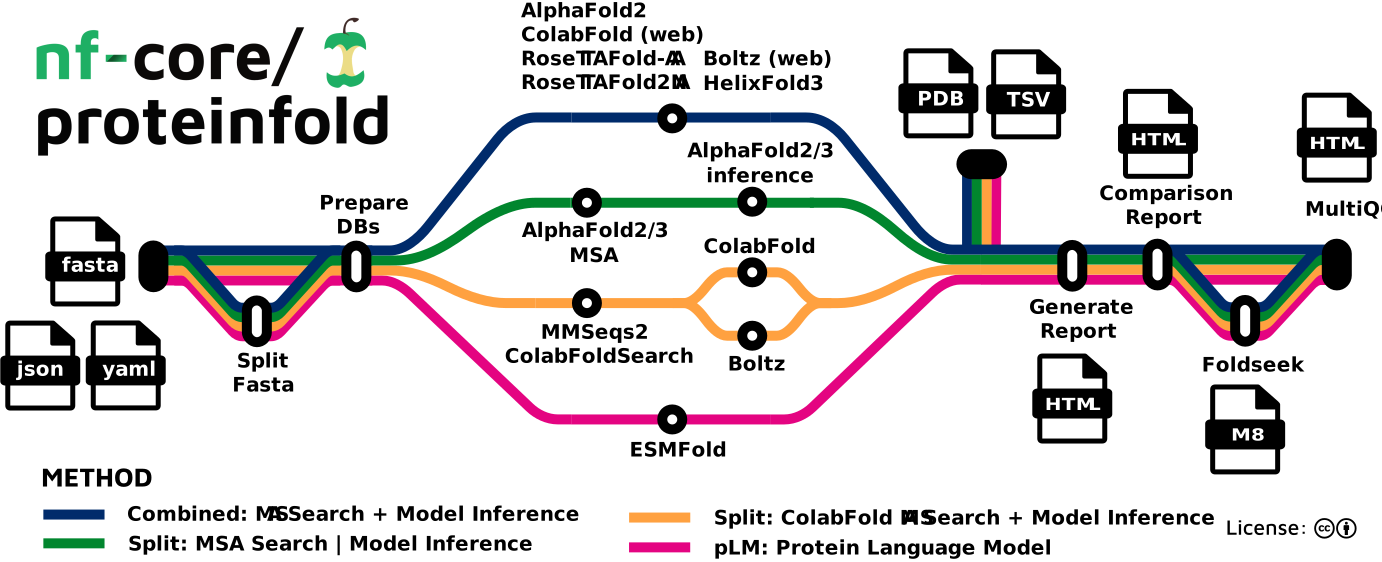
ESMFold
Boltz
ColabFold
MMSeqs2
ColabFoldSearch
AlphaFold2/3
MSA
AlphaFold2/3
inference
Boltz (web)
HelixFold3
AlphaFold2
ColabFold (web)
RoseT
T
AFold-A
A
RoseT
T
AFold2N
A
Split
Fasta
Prepare
DBs
fasta
json
yaml
PDB
TSV
Generate
Report
MultiQC
Comparison
Report
Combined: MS
A
Search + Model Inference
Split: MSA Search | Model Inference
Split: ColabFold MS
A
Search + Model Inference
pLM: Protein Language Model
HTM
L
HTM
L
HTM
L
Foldseek
M8
1
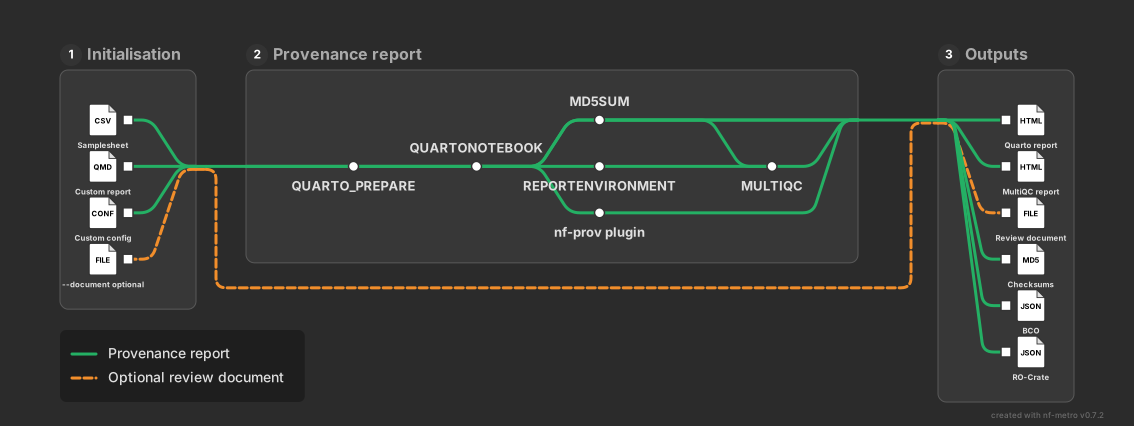
Initialisation
2
Provenance report
3
Outputs
CSV
Samplesheet
QMD
Custom report
CONF
Custom config
FILE
--document optional
HTML
Quarto report
HTML
MultiQC report
FILE
Review document
MD5
Checksums
JSON
BCO
JSON
RO-Crate
QUARTO_PREPARE
QUARTONOTEBOOK
MD5SUM
REPORTENVIRONMENT
nf-prov plugin
MULTIQC
Provenance report
Optional review document
created with nf-metro v0.7.2
Create ID map
cat
awk
Postprocess clusters
awk
Find near-identical sequences
Diamond
nf-
core/
reportho
Fasta
Sequence query
ID
Uniprot ID query
Identify sequence
OMA
Identify taxon
OMA
OMA
PANTHER
OrthoInspector
EggNOG
Online
Local
Online
Local
Online
Local
csv
csv
csv
csv
Single predictions
Query information
txt
Fetch ortholog predictions
Dump parameters
cat
Generate reports
React
Version 1.1.0
Reliable Rudder
Core workflow
Optional report generation
Core data flow
Report data flow
html
Per-query reports
Choose one
Subworkflow
html
Summary report
Generate summary
MultiQC
Split by source
Python
OMA
Uniprot
RefSeq
Ensembl
Get sequences
fasta
Ortholog sequences
Merge
cat
Split by taxon
awk
Merge IDs
Filter hits
Make score table
csvmerge
Python
csv
Score table
Python
Plot orthologs
ggplot
list
Filtered orthologs
Comparison plots
Fastq
Fastq
png
Python
Ortholog statistics
Calculate statistics
list
Score orthologs
1
Pre-processing
2
Genome alignment & quantification
3
Pseudo-alignment & quantification
4
Post-processing
5
Quality control & reporting
FASTQ
HTML
HTML
HTML
STAR
STAR
Picard
Picard
RSeQC
RSeQC
Bowtie2
Bowtie2
HISAT2
HISAT2
Salmon
Salmon
Kallisto
Kallisto
cat FASTQ
cat FASTQ
BEDTools
BEDTools
Preseq
Preseq
UMI-tools Dedup
UMI-tools Dedup
tximport
tximport
FastQC
FastQC
RSEM
RSEM
bedGraphToBigWig
bedGraphToBigWig
Qualimap
Qualimap
Salmon
Salmon
Sum. Exp.
Sum. Exp.
UMI-tools Extract
UMI-tools Extract
tximport
tximport
StringTie
StringTie
dupRadar
dupRadar
MultiQC
MultiQC
fastp
fastp
Sum. Exp.
Sum. Exp.
featureCounts
featureCounts
Trim Galore!
Trim Galore!
FastQC
FastQC
DESeq2 PCA
DESeq2 PCA
MultiQC
MultiQC
Kraken2/Bracken
Kraken2/Bracken
BBSplit
BBSplit
Sylph
Sylph
SortMeRNA
SortMeRNA
RiboDetector
RiboDetector
MultiQC
MultiQC
Bowtie2
Bowtie2
FastQC
FastQC
Infer Strand.
Infer Strand.
Aligner: STAR, Quantification: RSEM
Aligner: STAR, Quantification: Salmon (default)
Aligner: Bowtie2 (prokaryotic), Quantification: Salmon
Aligner: HISAT2, Quantification: None
Pseudo-aligner: Salmon, Quantification: Salmon
Pseudo-aligner: Kallisto, Quantification: Kallisto
created with nf-metro v2.1.0
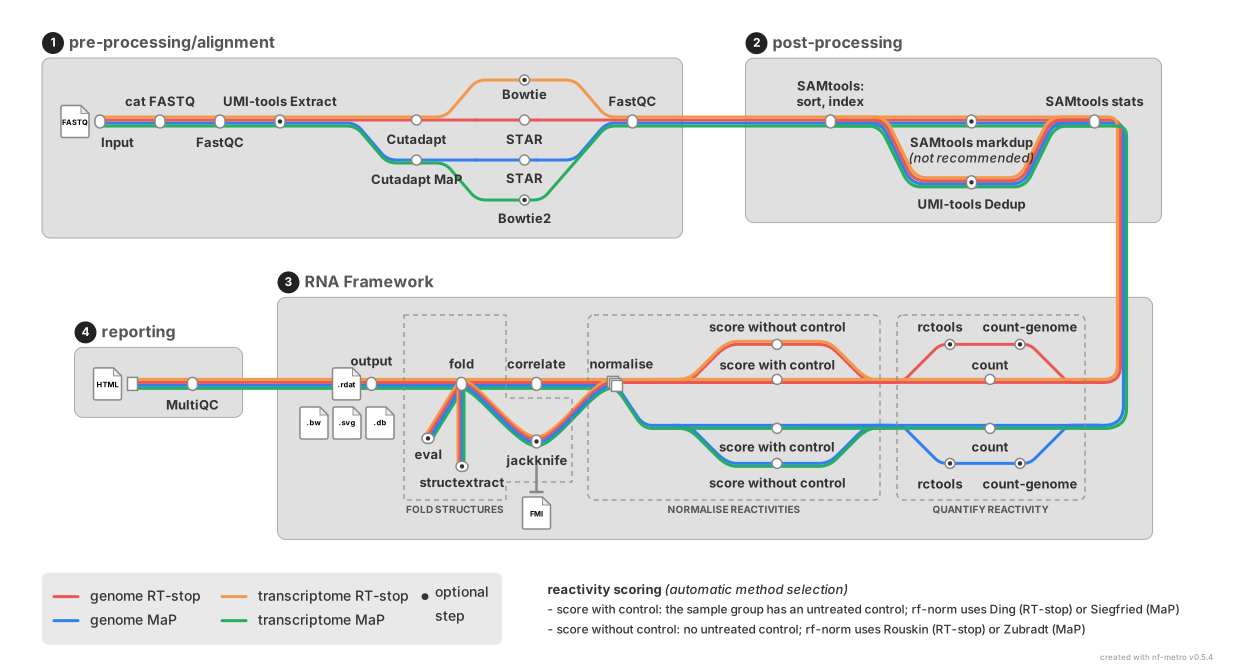
1
pre-processing/alignment
2
post-processing
3
RNA Framework
4
reporting
Bowtie
Bowtie2
FASTQ
Input
SAMtools:
sort, index
MultiQC
cat FASTQ
SAMtools markdup
(not recommended)
UMI-tools Dedup
FastQC
SAMtools stats
rctools
rctools
UMI-tools Extract
score with control
score with control
score without control
score without control
Cutadapt
Cutadapt MaP
normalise
STAR
correlate
STAR
FastQC
fold
output
genome RT-stop
genome MaP
transcriptome RT-stop
transcriptome MaP
.bw
.svg
.db
created with nf-metro v0.5.4
count-genome
count-genome
count
count
jackknife
FMI
eval
structextract
QUANTIFY REACTIVITY
NORMALISE REACTIVITIES
FOLD STRUCTURES
.rdat
HTML
optional
step
reactivity scoring
(automatic method selection)
-
score with control
: the sample group has an untreated control; rf-norm uses Ding (RT-stop) or Siegfried (MaP)
-
score without control
: no untreated control; rf-norm uses Rouskin (RT-stop) or Zubradt (MaP)
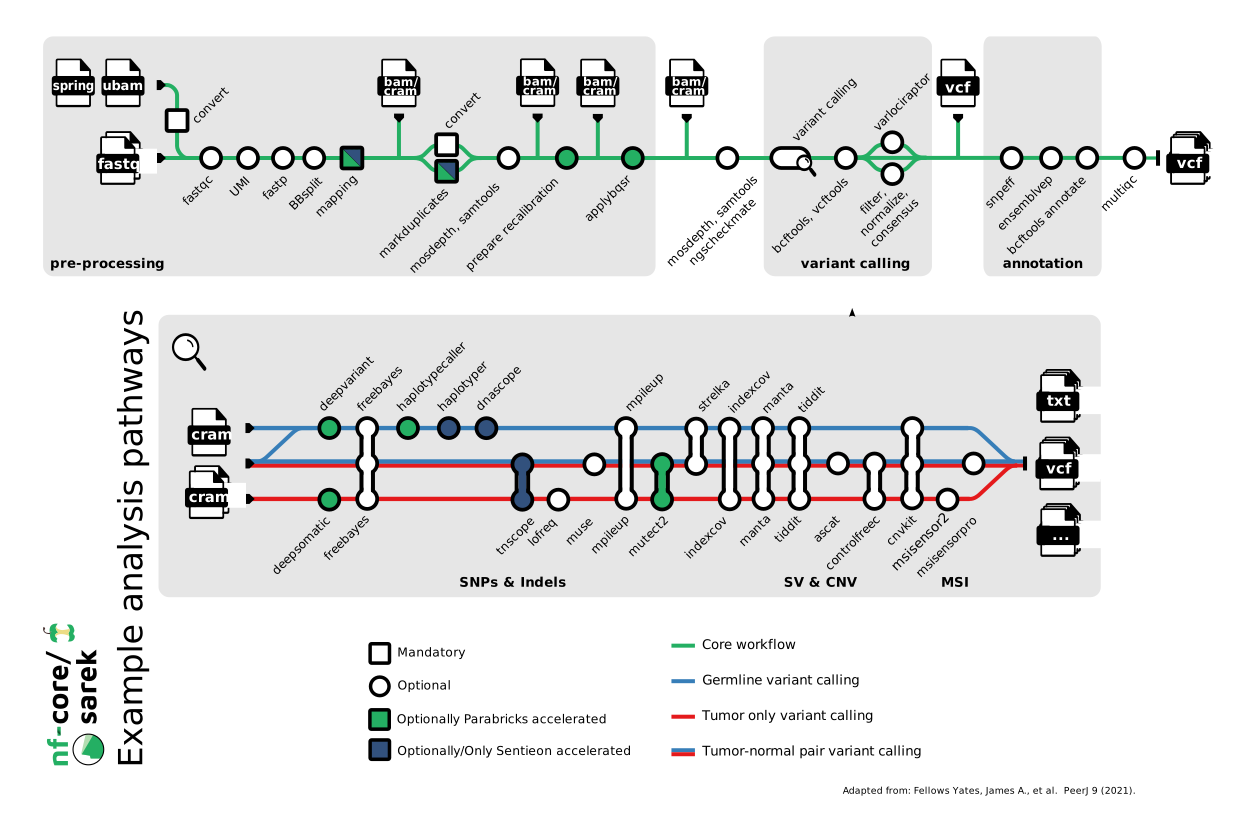
indexcov
indexcov
UMI
mapping
vcf
txt
...
fastqc
fastp
fastq
ensemblvep
snpeff
mosdepth, samtools
bcftools, vcftools
mosdepth, samtools
ngscheckmate
varlociraptor
multiqc
convert
bam/
cram
bam/
cram
bam/
cram
vcf
markduplicates
bam/
cram
variant calling
pre-processing
annotation
variant calling
Tumor-normal pair variant calling
Adapted from: Fellows Yates, James A., et al. PeerJ 9 (2021).
Core workflow
Germline variant calling
Tumor only variant calling
BBsplit
prepare recalibration
applybqsr
filter,
normalize,
consensus
bcftools annotate
manta
tiddit
tiddit
mutect2
ascat
msisensorpro
controlfreec
cnvkit
vcf
muse
manta
lofreq
tnscope
convert
Example analysis pathways
ubam
spring
mpileup
mpileup
haplotyper
dnascope
deepvariant
freebayes
haplotypecaller
cram
cram
deepsomatic
freebayes
Optional
Mandatory
Optionally/Only Sentieon accelerated
Optionally Parabricks accelerated
SNPs & Indels
SV & CNV
MSI
strelka
msisensor2
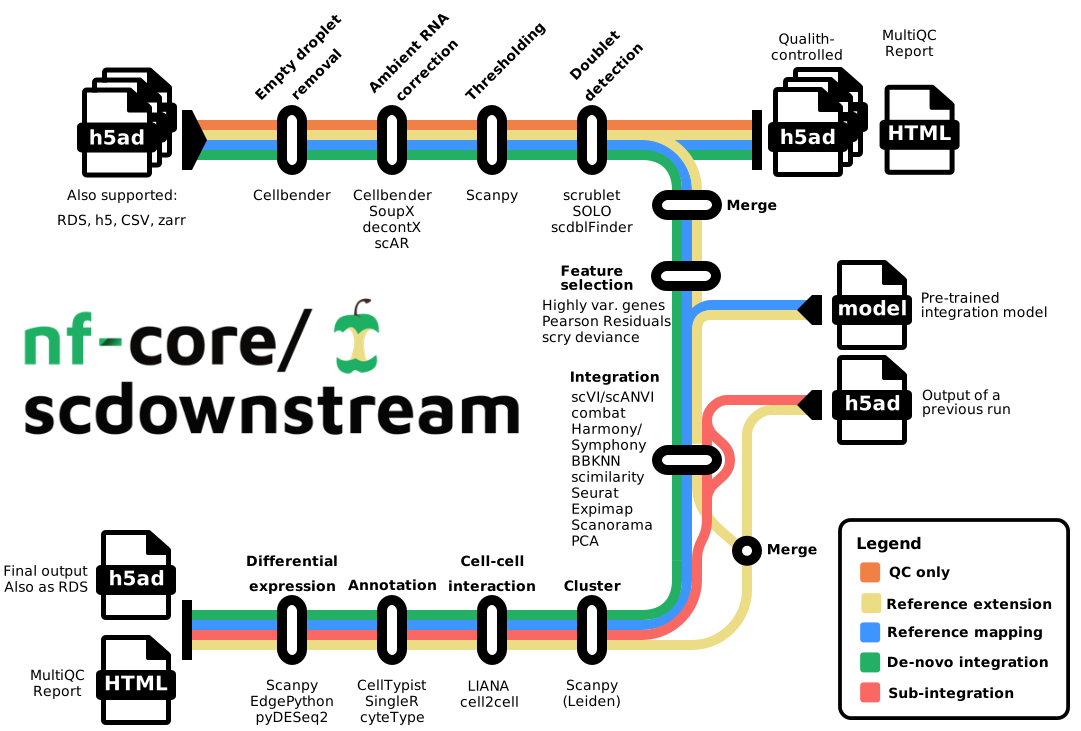
Legend
De-novo integration
QC only
Feature
selection
Merge
Merge
Also supported:
RDS, h5, CSV, zarr
Pre-trained
integration model
Output of a
previous run
Cellbender
Cellbender
SoupX
decontX
scAR
LIANA
cell2cell
Scanpy
(Leiden)
Scanpy
CellTypist
SingleR
cyteType
Scanpy
EdgePython
pyDESeq2
Final output
Also as RDS
MultiQC
Report
MultiQC
Report
Qualith-
controlled
scrublet
SOLO
scdblFinder
scVI/scANVI
combat
Harmony/
Symphony
BBKNN
scimilarity
Seurat
Expimap
Scanorama
PCA
Highly var. genes
Pearson Residuals
scry deviance
Empty droplet
removal
Ambient RNA
correction
Thresholding
Doublet
detection
Reference mapping
Reference extension
Sub-integration
Integration
Cluster
Cell-cell
interaction
Annotation
model
h5ad
h5ad
HTML
HTML
fastq
fastq
h5ad
fastq
fastq
h5ad
Differential
expression
1
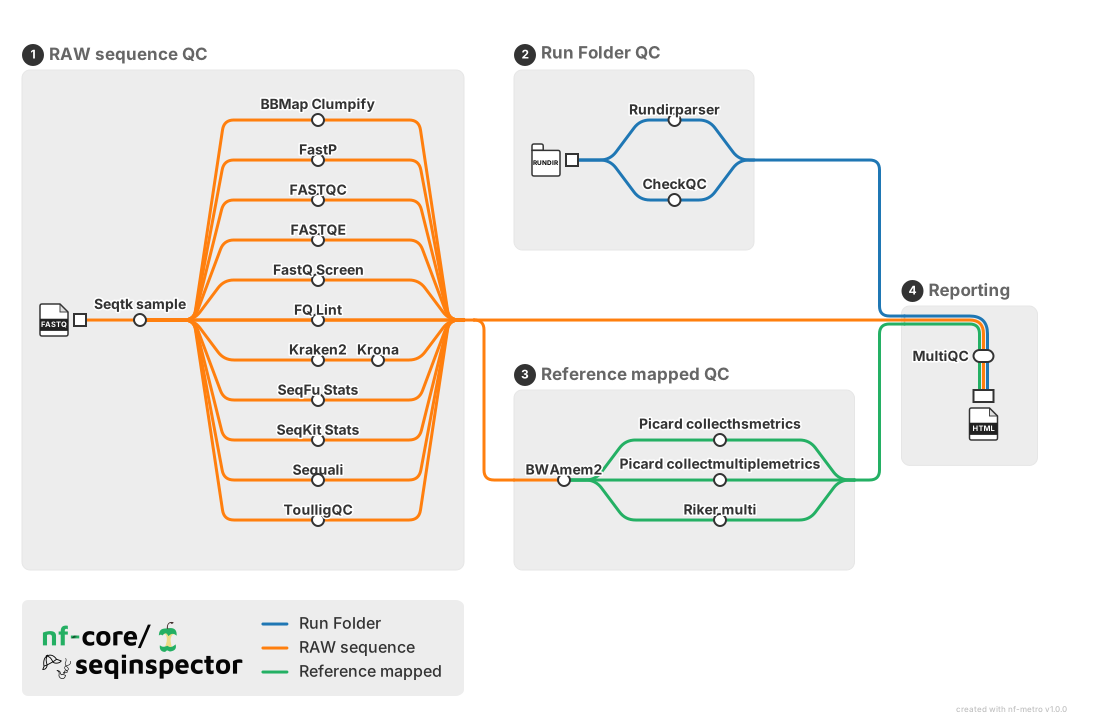
RAW sequence QC
3
Reference mapped QC
4
Reporting
FASTQ
HTML
MultiQC
MultiQC
BWAmem2
BWAmem2
2
Run Folder QC
RUNDIR
CheckQC
CheckQC
Rundirparser
Rundirparser
Picard collecthsmetrics
Picard collecthsmetrics
Seqtk sample
Seqtk sample
Picard collectmultiplemetrics
Picard collectmultiplemetrics
Riker multi
Riker multi
BBMap Clumpify
BBMap Clumpify
FastP
FastP
FASTQC
FASTQC
FASTQE
FASTQE
FastQ Screen
FastQ Screen
FQ Lint
FQ Lint
Kraken2
Kraken2
SeqFu Stats
SeqFu Stats
SeqKit Stats
SeqKit Stats
Sequali
Sequali
ToulligQC
ToulligQC
Krona
Krona
Run Folder
RAW sequence
Reference mapped
created with nf-metro v1.0.0
1
Read preparation
2
miRNA and isomiR analysis
3
Genome analysis
4
Reporting
FASTQ
Small RNA reads
HTML
QC report
Bowtie
Bowtie
miRTrace
miRTrace
mirtop
mirtop
Bowtie
Bowtie
miRDeep2
miRDeep2
FastQC
FastQC
edgeR
edgeR
MultiQC
MultiQC
UMI-tools
UMI-tools
fastp
fastp
UMICollapse
UMICollapse
FastQC
FastQC
Bowtie2
Bowtie2
miRNA quantification
Quality control
isomiR quantification
Genome quantification (optional)
miRNA discovery (optional)
created with nf-metro v2.1.0
txt
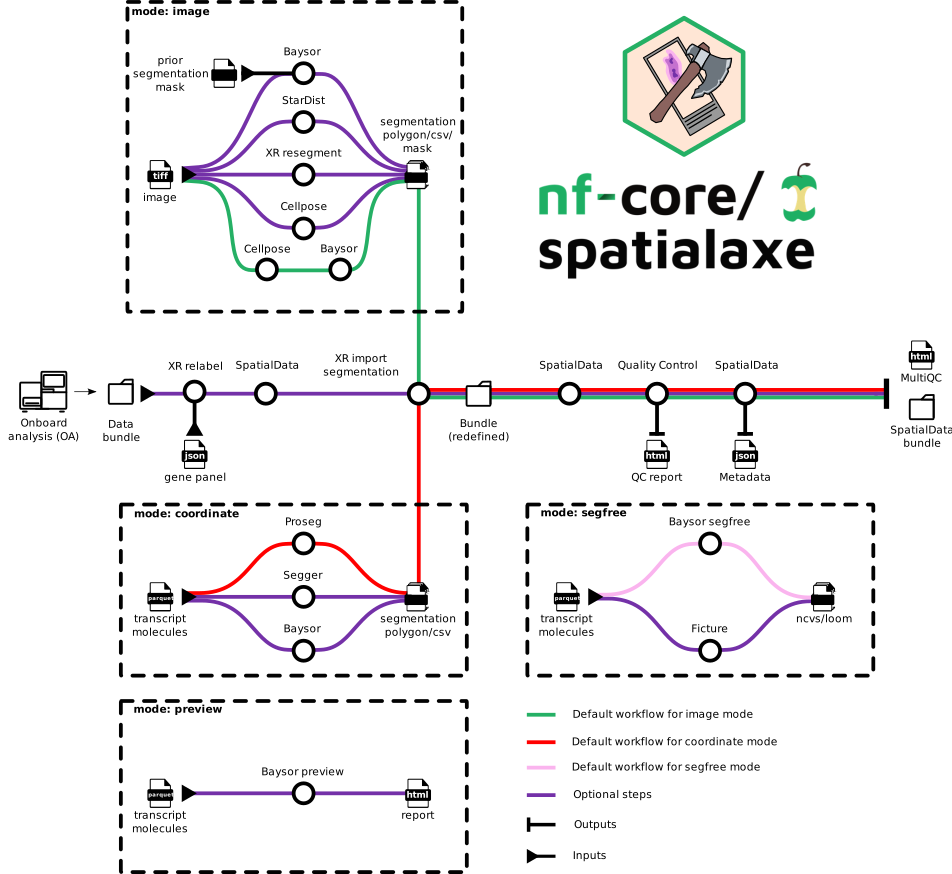
Xenium onboard
analysis (XOA)
vcf
XR import
segmentation
Data
bundle
html
QC report
json
Metadata
json
gene panel
Bundle
(redefined)
html
MultiQC
SpatialData
bundle
XR relabel
SpatialData
SpatialData
SpatialData
Quality Control
Onboard
analysis (OA)
Default workflow for image mode
Default workflow for coordinate mode
Default workflow for segfree mode
Optional steps
Outputs
Inputs
mode: image
XR resegment
StarDist
Cellpose
Baysor
Cellpose
tiff
image
Baysor
prior
segmentation
mask
segmentation
polygon/csv
segmentation
polygon/csv/
mask
Segger
Proseg
Baysor
parquet
transcript
molecules
report
Baysor preview
html
parquet
transcript
molecules
mode: coordinate
ncvs/loom
Baysor segfree
Ficture
parquet
transcript
molecules
mode: segfree
mode: preview
mode: coordinate
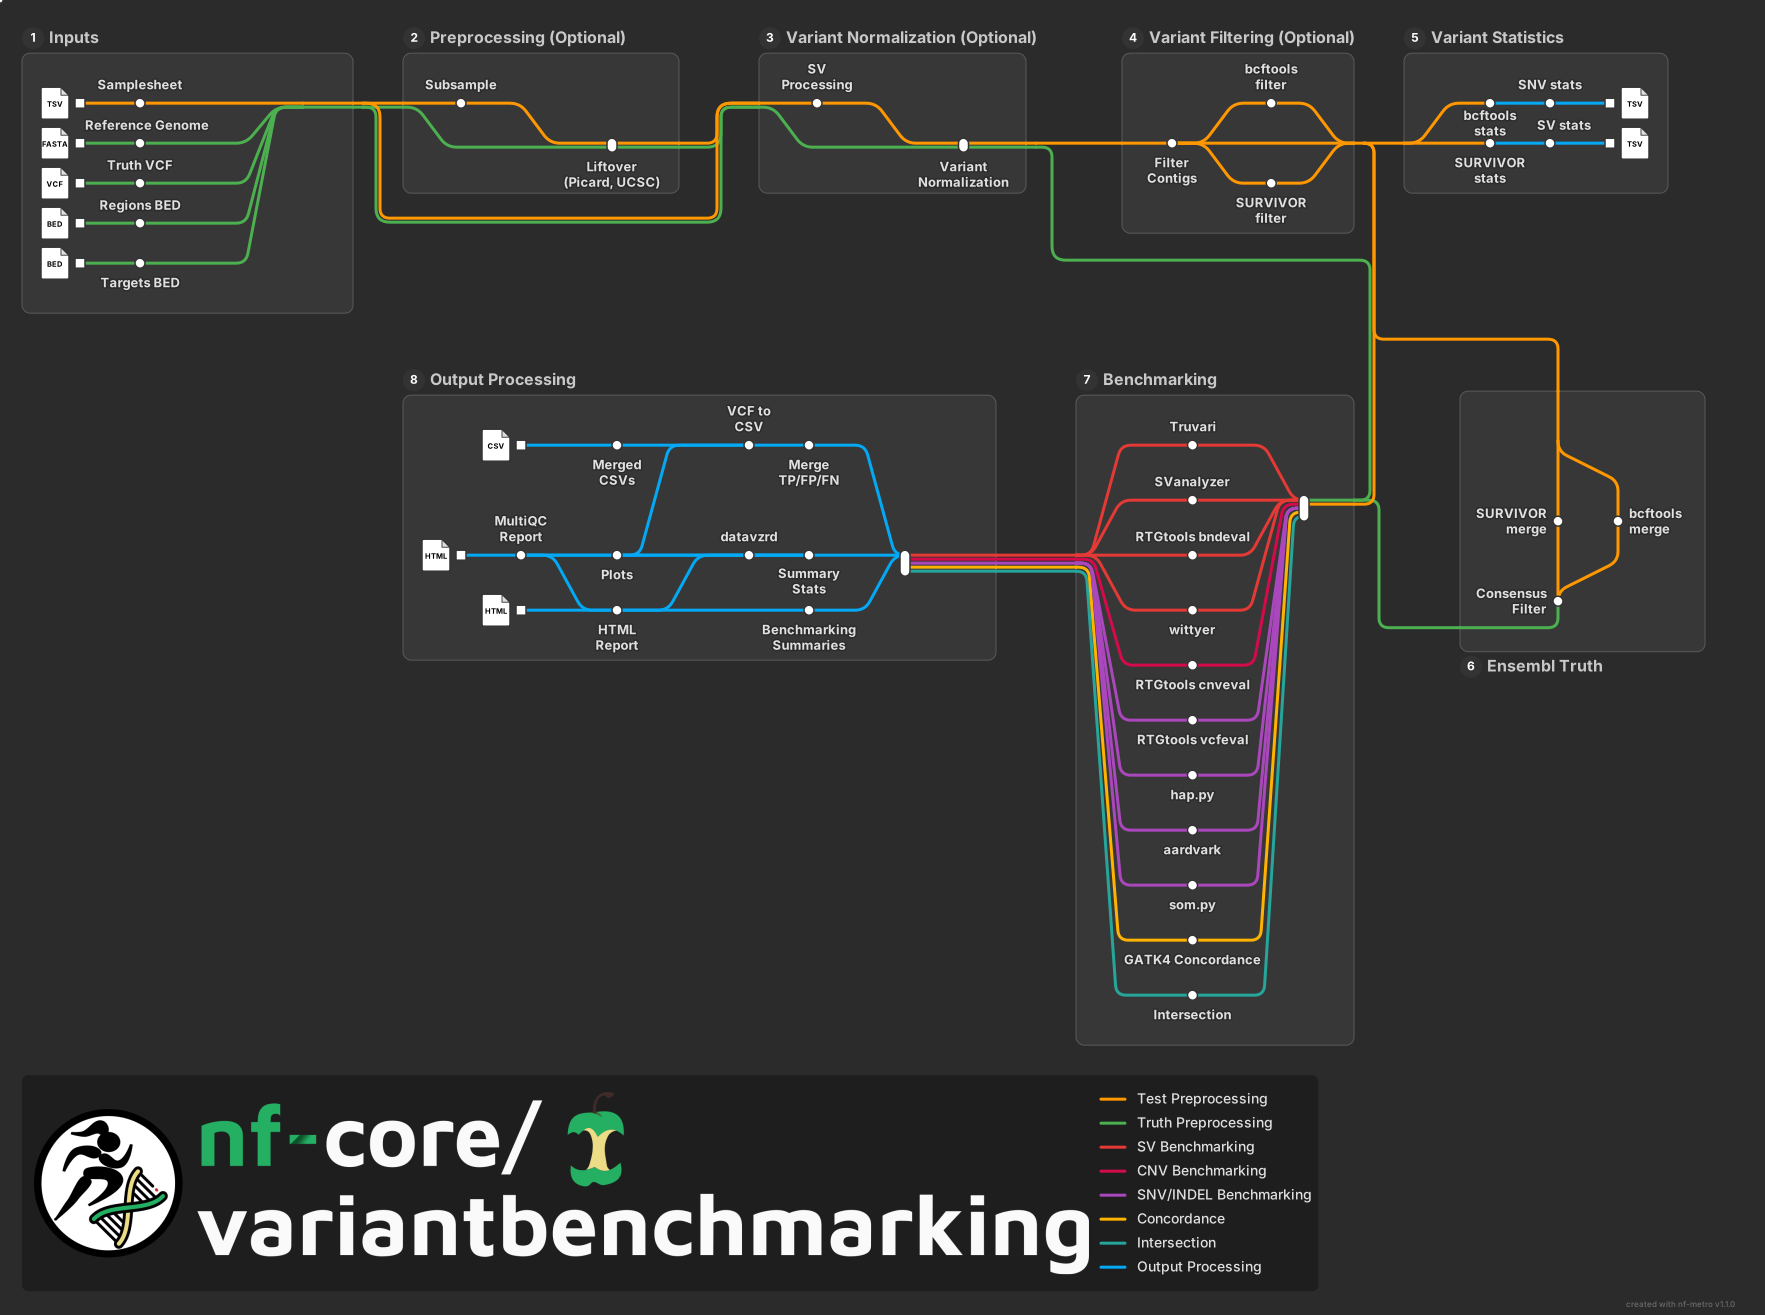
1
Inputs
2
Preprocessing (Optional)
3
Variant Normalization (Optional)
4
Variant Filtering (Optional)
6
Ensembl Truth
7
Benchmarking
8
Output Processing
5
Variant Statistics
FASTA
VCF
BED
BED
TSV
CSV
HTML
HTML
TSV
TSV
Subsample
Subsample
SV
Processing
SV
Processing
Filter
Contigs
Filter
Contigs
bcftools
stats
bcftools
stats
SURVIVOR
stats
SURVIVOR
stats
bcftools
filter
bcftools
filter
Samplesheet
Samplesheet
SURVIVOR
merge
SURVIVOR
merge
Truvari
Truvari
SURVIVOR
filter
SURVIVOR
filter
Reference Genome
Reference Genome
Liftover
(Picard, UCSC)
Liftover
(Picard, UCSC)
Variant
Normalization
Variant
Normalization
bcftools
merge
bcftools
merge
SVanalyzer
SVanalyzer
Truth VCF
Truth VCF
RTGtools bndeval
RTGtools bndeval
Regions BED
Regions BED
wittyer
wittyer
Targets BED
Targets BED
RTGtools cnveval
RTGtools cnveval
RTGtools vcfeval
RTGtools vcfeval
Merge
TP/FP/FN
Merge
TP/FP/FN
SNV stats
SNV stats
hap.py
hap.py
Summary
Stats
Summary
Stats
aardvark
aardvark
SV stats
SV stats
Benchmarking
Summaries
Benchmarking
Summaries
som.py
som.py
GATK4 Concordance
GATK4 Concordance
Intersection
Intersection
Consensus
Filter
Consensus
Filter
VCF to
CSV
VCF to
CSV
datavzrd
datavzrd
Merged
CSVs
Merged
CSVs
Plots
Plots
HTML
Report
HTML
Report
MultiQC
Report
MultiQC
Report
Test Preprocessing
Truth Preprocessing
SV Benchmarking
CNV Benchmarking
SNV/INDEL Benchmarking
Concordance
Intersection
Output Processing
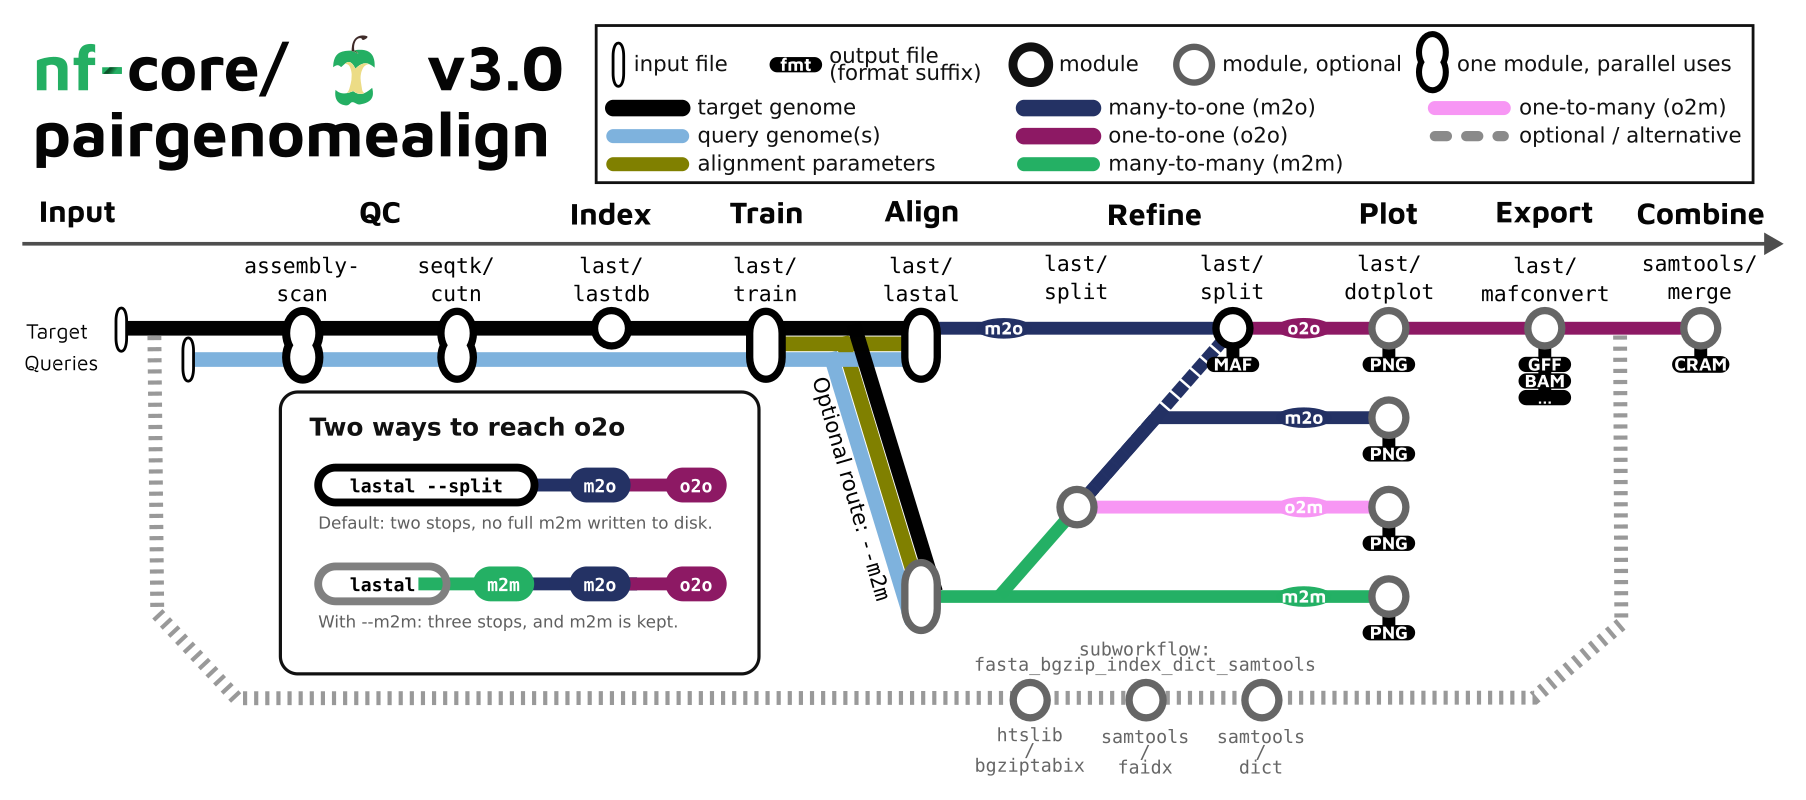
created with nf-metro v1.1.0